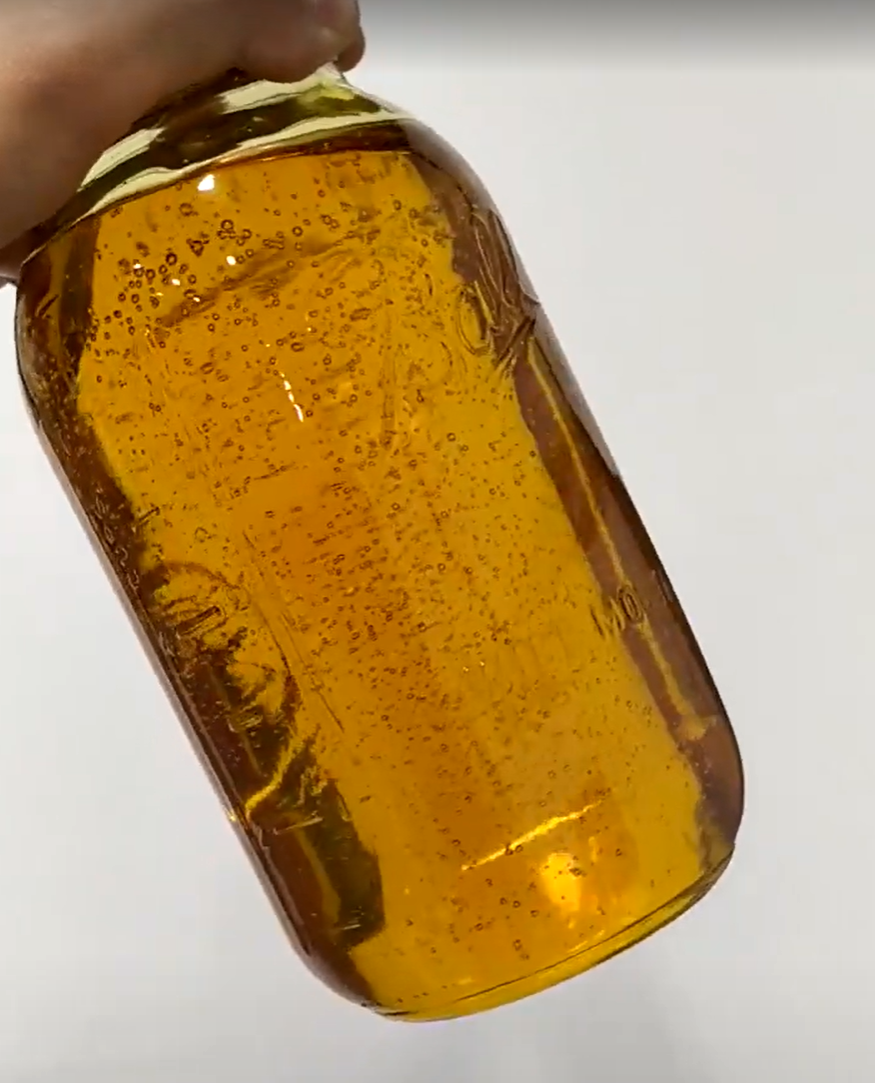
Devolatilization is often defined as the removal of volatile compounds within a substance. This is often necessary to further refine abotanical extractviadistillation. Devolatilization is anevaporative process that drives volatiles from abotanical extractwhen heat is applied. Typically this is a separate process performed prior todistillation for the sake of efficiency.
Devolatilization as a process is used to remove volatile compounds from abotanical extract, convert compounds into a more distillable form, and ensure an extract is effectivelypurged of residualsolvent prior to performingdistillation. When abotanical extractis properly devolatilized, it results in a smoother, more efficientdistillation of abotanical extract.
One of the more common forms ofdevolatilization of abotanical extractisdecarboxylation.Decarboxylation is a chemical reaction that removes a carboxyl group from an organic compound releasing carbon dioxide. Typicallydecarboxylation is performed to remove loosely bonded carboxylic acids from the atomic chain of a botanical compound converting it into a more stable, easily distilled form. The most common use fordecarboxylation is to convert acidic phytocannabinoids into their non-acidic active forms byevaporating the loosely bonded carboxyl group from the compound resulting in the release of CO2.
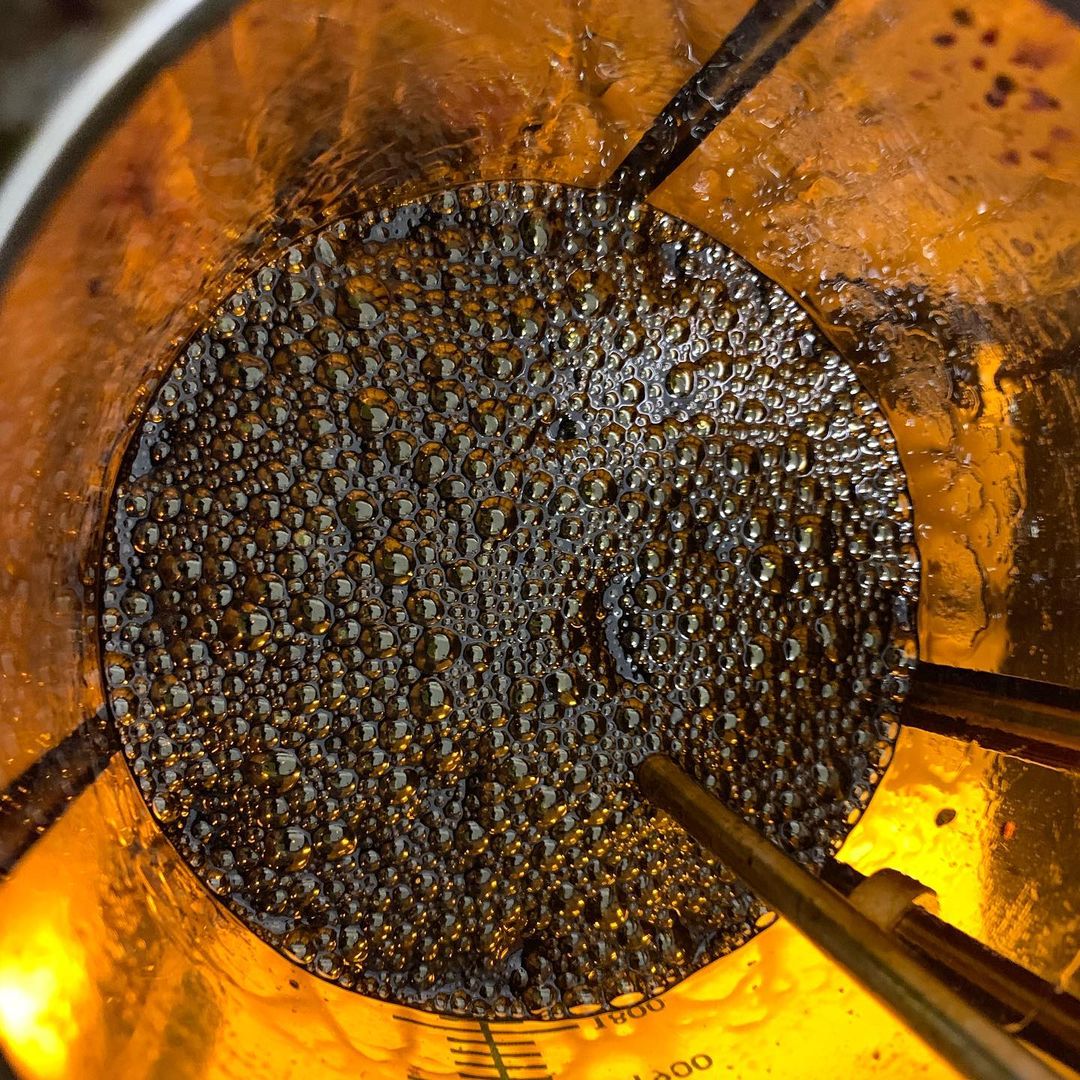
Begining ofDevolatilizaiton
In this case,decarboxylation is necessary prior todistillation because thedecarboxylation process occurs before theevaporation of the cannabinoid when heat is applied to the extract. This makes decarboxylation necessary to refine a cannabinoid-rich extract further into adistillate.
Both devolatilization and decarboxylation occur over time with the application of heat. The rate at which devolatilization is performed depends on how much heat is applied to the extract over time. While this process can be achieved quickly at high heat, this may evaporate some of the lower boiling point target compounds found within the matrix of a botanical extract. This is why most devolatilization processes call for moderate heat just below the lowest boiling point of the target compounds the operator is looking to distill. This results in the evaporation of all the lower boiling point compounds within the botanical extract, prepping it for an efficient distillation of the target compounds.
While this process can be performed within ashort path distillation system prior todistillation, it is best to perform this process in a separate apparatus for processing efficiency and not allow volatiles to contaminate thevacuumpump of theshort path distillationapparatus.
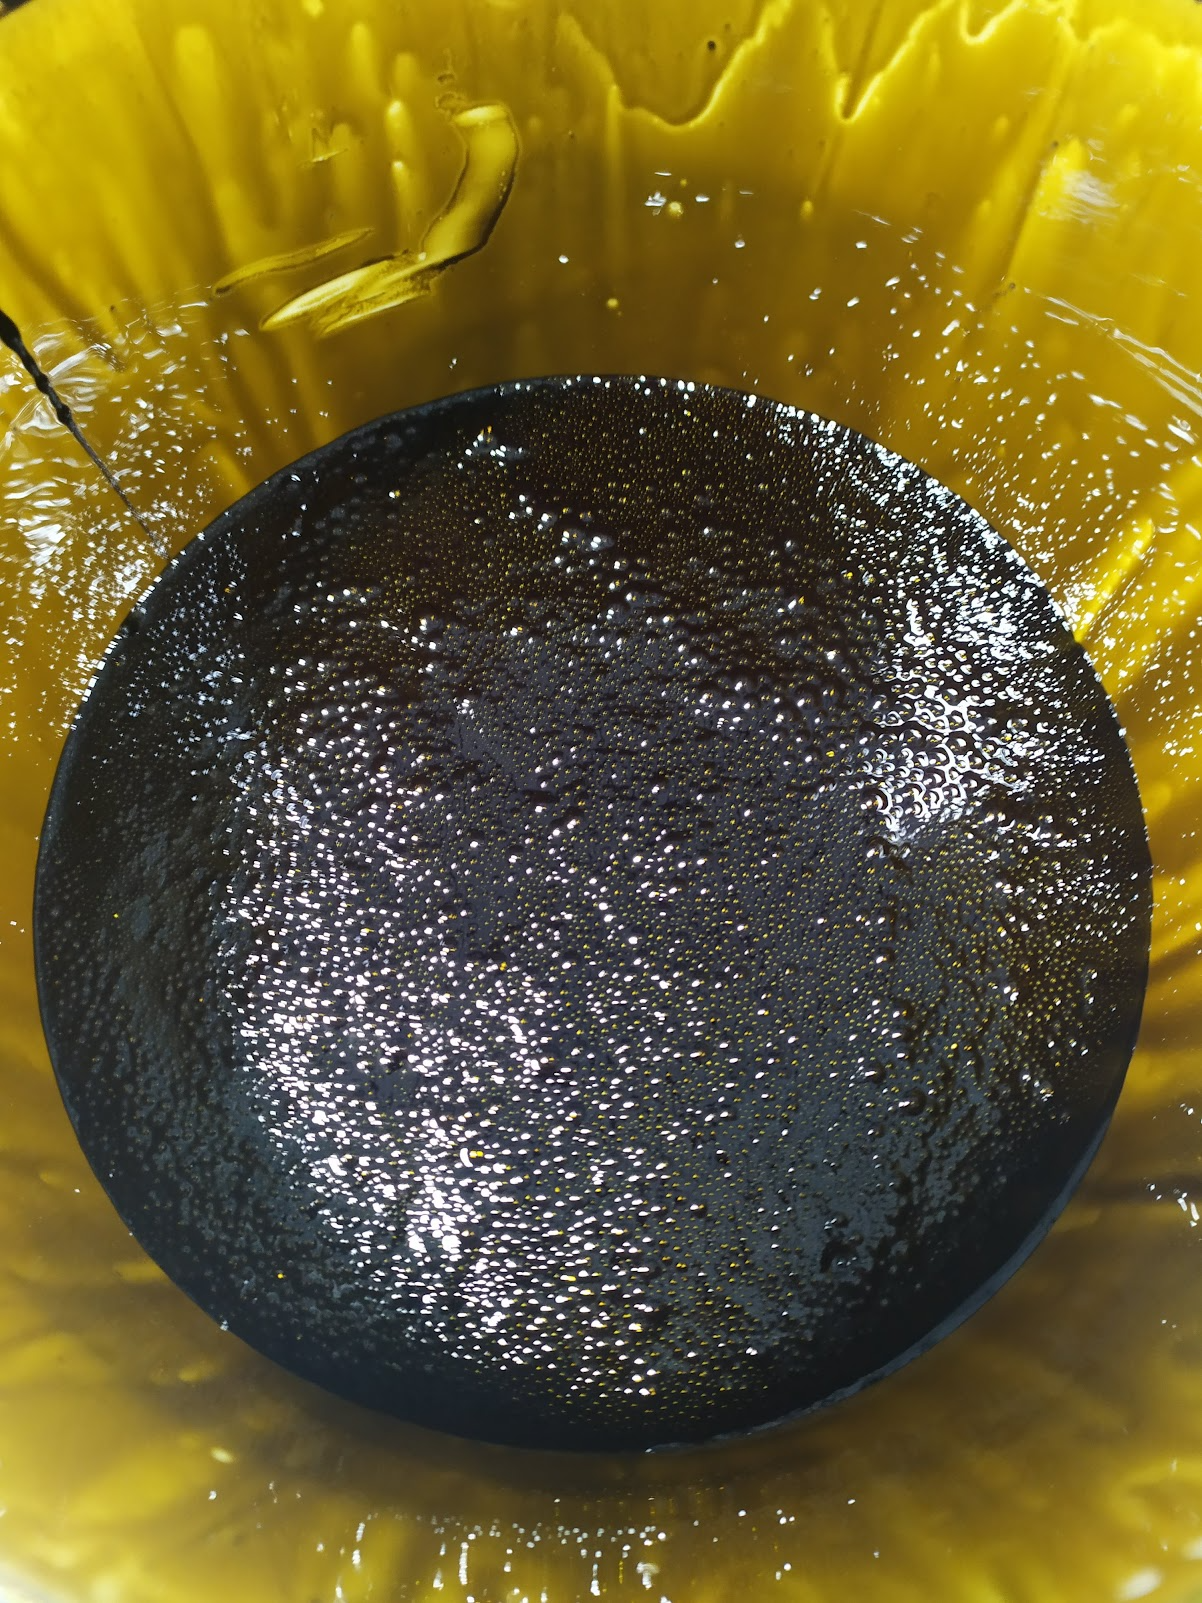
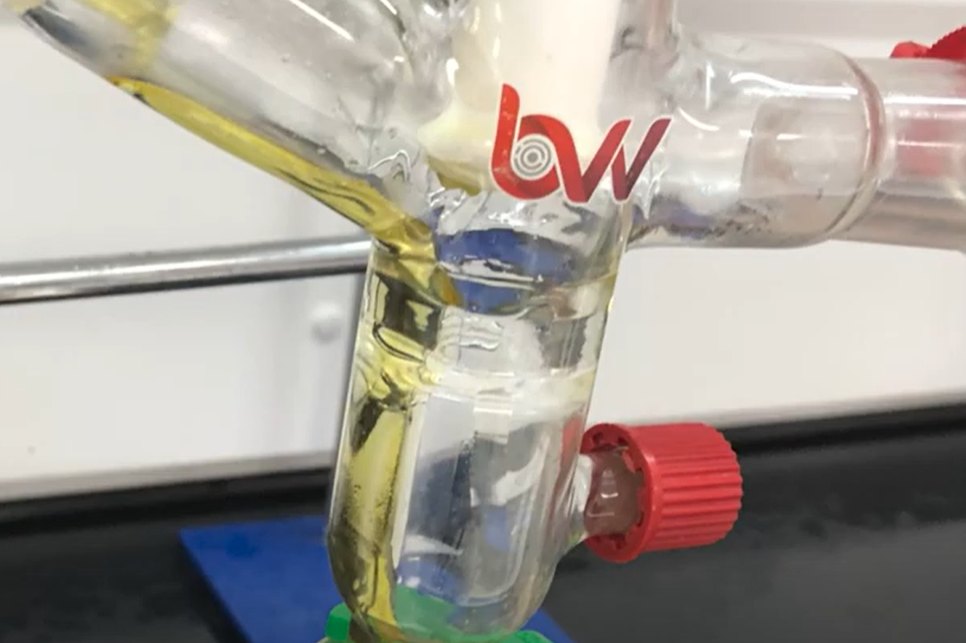
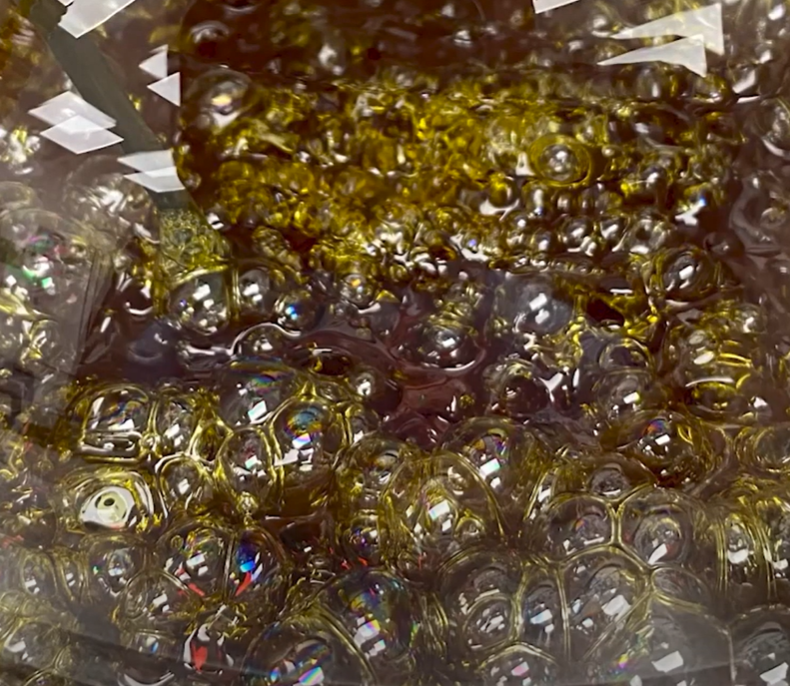
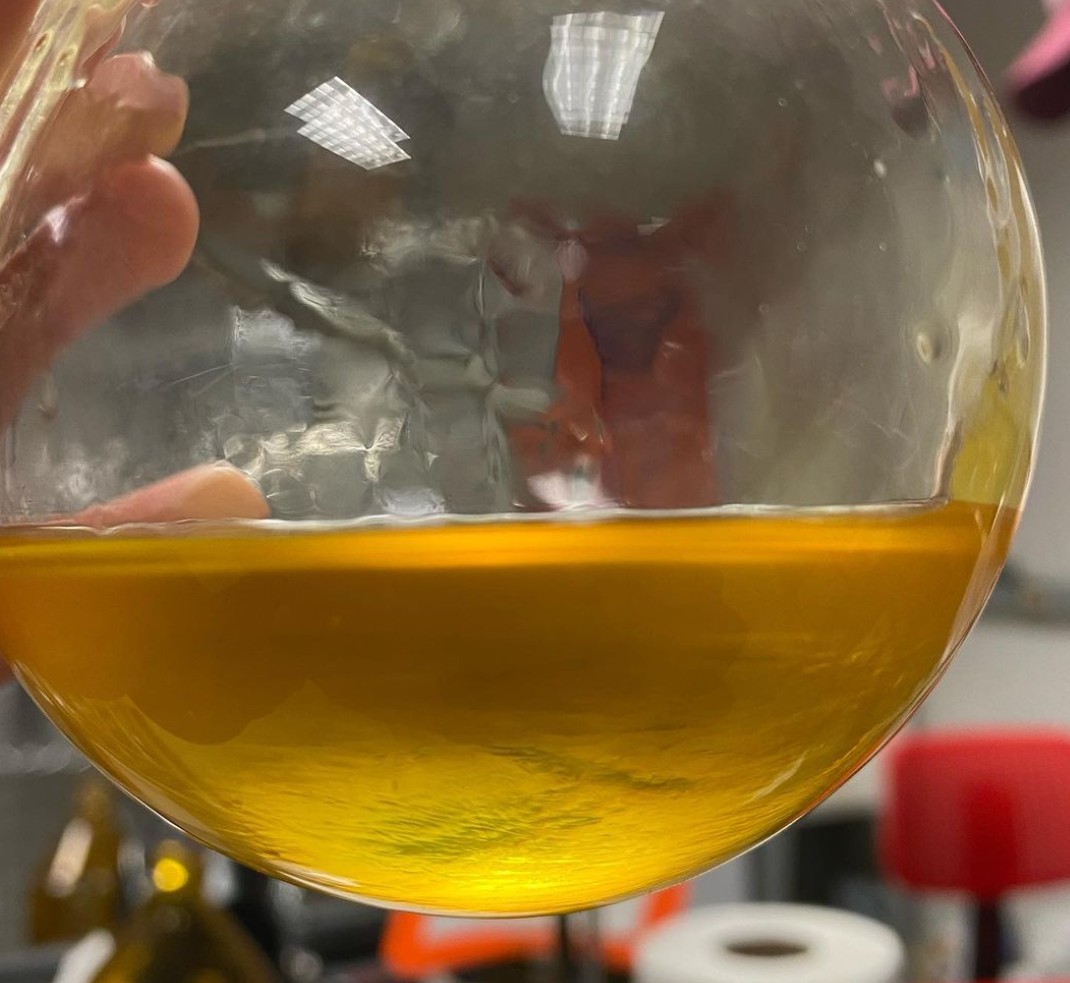
Partially Devolatilized Oil
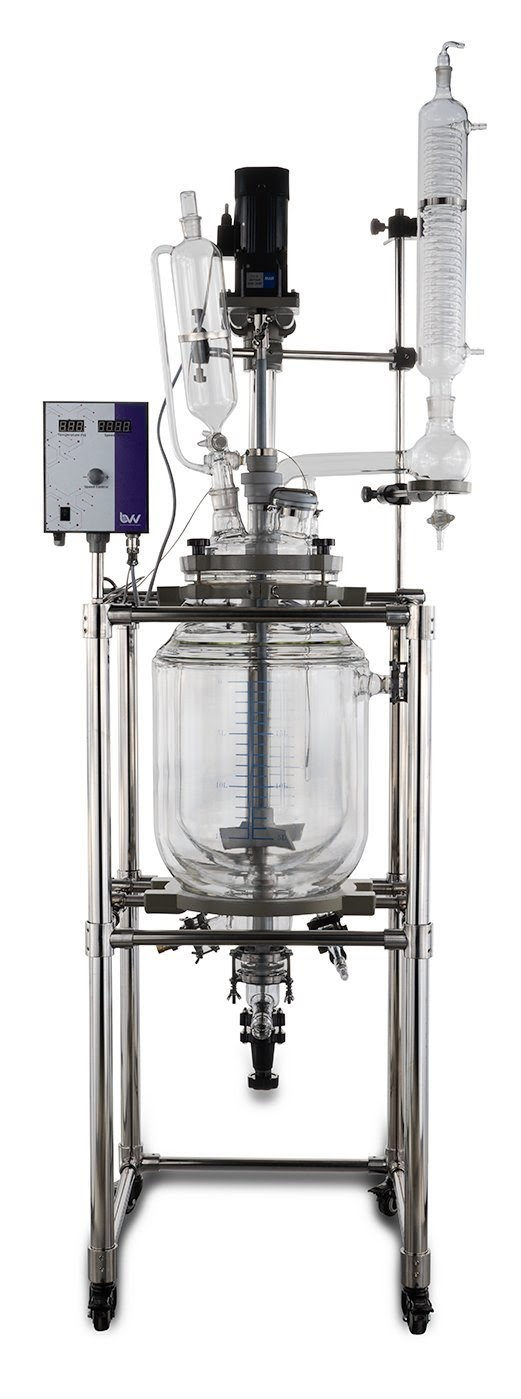
Devolatilization of a botanical extractis typically performed inside astainless steelorglassreactorby heating the extract just below the lowestboiling pointof the target compounds within the botanical extract. While this can be accomplished at ambient pressures,devolatilization can be achieved more efficiently undervacuum with agitation.Devolatilization is typically performed by incrementally heating an extract undervacuum to the desireddevolatilization temperature until no more volatilesevaporate from thesolution. Whiledevolatilizationtemperature is highly variable based on the target compounds of thedistillation, they usually range between 120C to 150C / 248F to 302F. A good indication that a botanical extractis fully devolatilized is when extract consistency stabilizes, and boiling is no longer observed at the setdevolatilization temperature. Onceevaporationhas ceased thedevolatilization process is complete. The extract should be allowed to cool to 65C/149F before transferring the devolatilized extract to another vessel for further processing.
Devolitilization andDecarboxylation should always be performed prior todistillation, not only for the sake of processing efficiency but also for safety. When an extract that has not been decarboxylated or devolatilized is loaded into adistillation system and heat andvacuum are applied the extract tends to expand rapidly as CO2 releases causing a violent reaction. This can result in excessive boiling that can overflow and contaminate the system, the contamination ofvacuum pump oil with volatiles, and the potential of pressure build-up resulting in an explosion of glass. For these reasons, it is always best to performdevolatilization in a separate apparatus. Thedevolatilizationapparatuses can range from a Jacketedreactor, heated, stirring plate under a fume hood, or convection oven with appropriate venting.
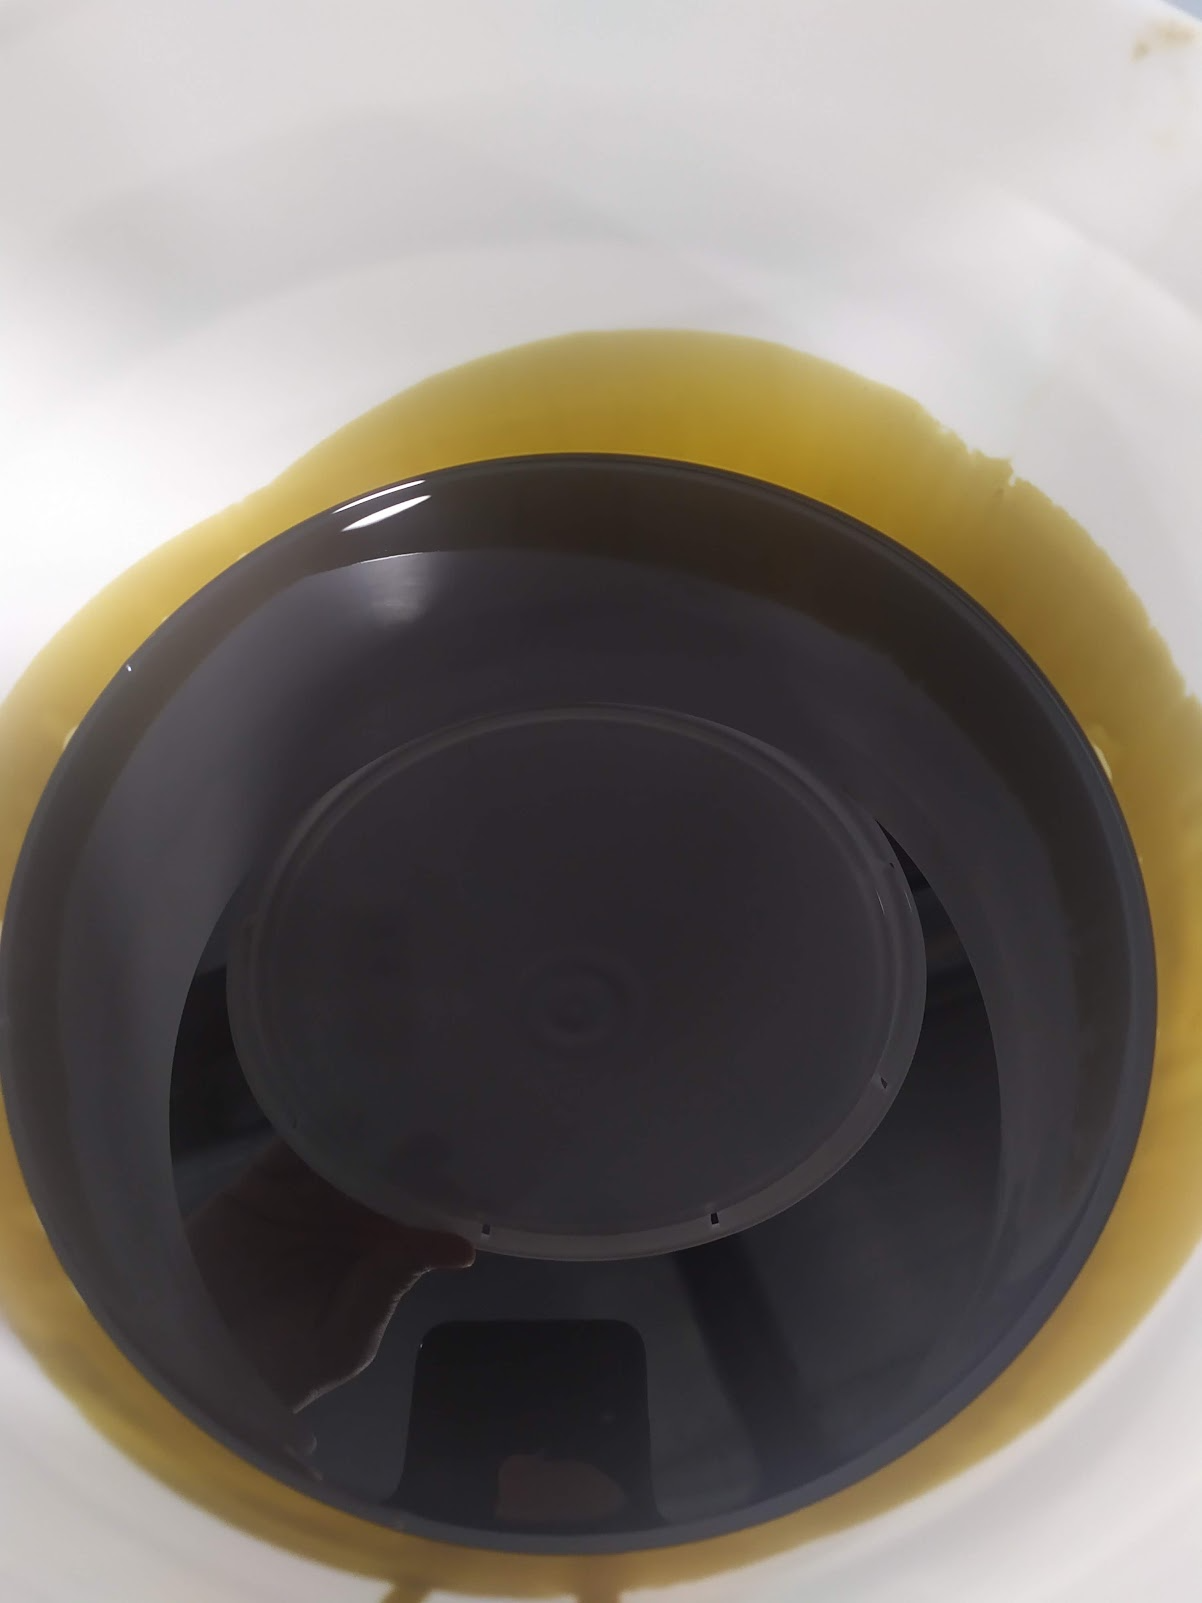
Full DecarboxylatedCrude Oil
DEVOLATILIZATION STANDARD OPERATING PROCEDURE
Purpose
The purpose of this procedure is to provide detailed instructions for thedevolatilization of abotanical extractprior todistillation.
Scope
This procedure applies to all lab technicians tasked with botanical extractdevolatilization.
Definitions/Acronyms
Personal Protection Equipment (PPE)Items worn to protect employees from exposure to hazardous materials and prevention of injury.
Safety Data Sheet (SDS)Provides useful information on chemicals, describing the hazards the chemical presents, and giving information on handling, storage, and emergency measures in case of an accident.
Devolatilization The removal of volatile compounds within a substance.
Decarboxylation A chemical reaction that removes a carboxyl group from a compound releasing CO2.
Safety
SDS Sheets: Seeextraction solvent SDS for detailed risks
PPE: The following should be worn by all lab personnel while winterizingbotanical extracts:
Protective eyewear
Lab coat
Gloves
5.Hazard Identification
Preparation and Use:
Abotanical extractwill be heated to remove any volatiles remaining in the extract prior todistillation.
Concentration- >10% residualextraction solvent.
Quantity- Residualextraction solvent.
Frequency- No newsolvent introduced.
Location- Jacketedreactor, headed stirring plate under a fume hood, or convection oven with appropriate venting.
Potential Hazards and Risks
Seeextraction solvent SDS for detailed risks.
The explosion from excessive internal pressure: thedevolatilization could explode if the internal pressure produced by off-gassed volatiles becomes too great.
Burns: Do not touch heated surfaces flask during operation.
Brokenglassware: Cracks or chips inglassware can result in glass rupturing causing chemical exposure and explosion hazard to the user. Always inspectglassware prior to use
6. Procedure
1. Transfer winterized andsolvent recoveredbotanical extractinto adevolatilization vessel ensuring extract volume does not exceed ⅔ the volume of thedevolatilization vessel.
2. Apply incremental heat until a temperature just below the lowestboiling pointof the target compounds you are preparing to distill is reached.
3. Continue to apply heat to the extract just below the temperature of your target compound's lowestboiling pointuntil extract consistency stabilizes and boiling is no longer observed.
4. Allow yoursolution to cool to 65C/149F before transferring the extract to a new vessel.
DISTILLATION
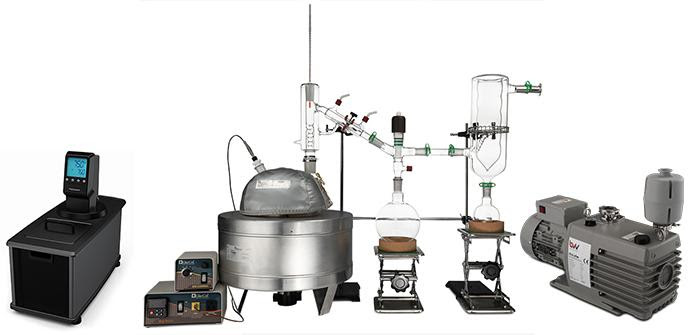
5L NeocisionShort PathDistillation Turnkey Kit By BVV
Distillation is a powerful purification technique that can drastically increase the purity of a botanical extract.Distillation can be simplified as the act of purifying a liquid by using selectiveevaporation and condensation. This purification process is implemented by heating the input material until target compounds areevaporated and then pulling the resultingvapor through a chilled condenser. It is condensed back into liquid form and collected.Distillation carries excellent utility in separating compounds in a mixture when there is a great enough difference between their boiling points and or theirvapor pressure.
Vapor pressureor equilibriumvapor pressureis the pressure of avapor in thermodynamic equilibrium within its condensed phase inside a closed container. When a liquid is confined to a closed container, an equilibrium exists between its liquid and itsgaseous phase. This equilibrium exists regardless of the temperature inside the container or the liquid because some particles within the liquid, essentially at any temperature, will always have enough energy to escape the intrinsic cohesive forces and enter thegaseous state.
When distilling an impure mixture of compounds like that found within a botanical extract, the various constituents contribute to the overall vapor pressure of the liquid depending on the concentration of compounds within the solution and their respective boiling points. The vapor pressure of a solution is affected by temperature alone and is not a function of volume or surface area. As the temperature of a solution increases or decreases, so does its vapor pressure. During the distillation process, when the temperature and the vapor pressure of a compound have increased past the ambient pressure of the closed system it is within, a compound will transform from a condensed state to a gaseous state referred to as evaporation.
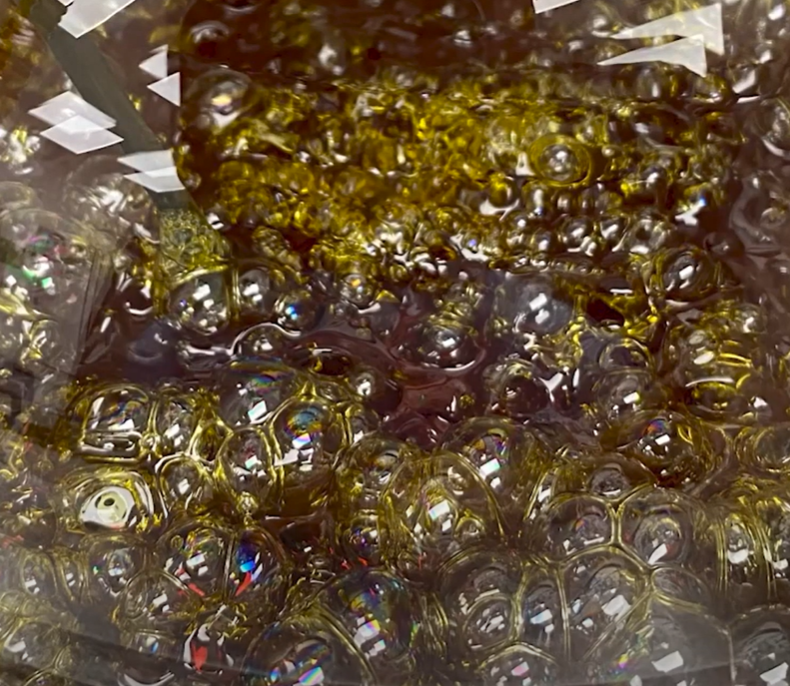
Evaporation Occurring
Equilibriumvapor pressure is an indication of a liquid'sevaporation rate. It relates to the tendency of particles to escape from their condensed phase into thevapor phase, which can be observed as boiling. When a liquid is boiling, its vapor pressureis equal to or greater than the ambient pressure within the closed system containing it causing compounds toevaporate from their condensed state. The rate at which these compounds areevaporated is determined by its vapor pressure which is a function of the temperature exerted upon its liquified state and the ambient pressure of the closed system that it must overcome.
Theevaporationof compounds happens at different rates depending on thevapor pressureof each constituent. Compounds with the highest vapor pressure tend toevaporate from thesolution first as heat is applied duringdistillation. Compoundsevaporatefrom thesolution in order based on their atmosphericboiling pointas heat is applied. Compounds with similarvapor pressuresare commonlyevaporated and condensed at the same temperature range making it challenging toisolate compounds in some instances completely.
Typically compounds within the same temperature range areevaporated, condensed, and collected into what is referred to as a fraction. These fractions are typically divided into three categories: the heads fraction containing the compounds thatevaporate before the target compounds, the main body containing the desired target compounds, and the tails fraction containing the compounds thatevaporate after the target compounds.
When it comes toevaporation, for a compound to undergo thephase changefrom a liquid to agas, it needs to overcome the ambient pressure within the system. Think of ambient pressure as an omnipresent weight pushing down at all times. The greater the ambient pressure the more energy in the form of heat is needed for compounds toevaporate.
This can be observed when boiling water at different elevations or atmospheric pressures. Atmospheric pressure is the weight of the atmosphere exerted on the earth's surface. Typically measured with a barometer, at sea level where atmospheric pressure is greatest, barometric pressure is about 14.7 PSI under standard conditions. There is less atmospheric pressure at higher elevations, resulting in reduced resistance toevaporation. As a result, water is observed to boil at a lower temperature at higher elevations. It requires more energy in the form of heat in order to boil at sea level due to the increased atmospheric pressure.
By reducing the amount of ambient pressure exerted upon a compound we can drastically reduce the amount of resistance a compound must overcome toevaporate. While energy in the form of heat is necessary to triggerevaporation by reducing the pressure within the system by pulling it under avacuum, the amount of energy in the form of heat necessary to induceevaporation is reduced. The most efficient way to reduce pressure within a closed system is to pull the system undervacuum with avacuum pump.
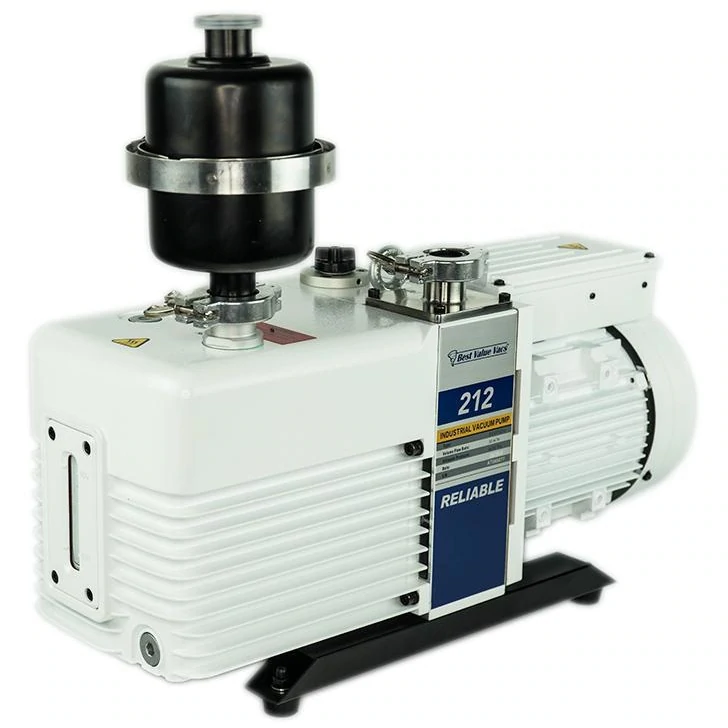
21.2CFM Pro Series Corrosion Resistant Two StageVacuum Pump By BVV
Vacuum can be considered negative air pressure or pressure measuring below atmospheric pressure. Air pressure is defined as atmospheric pressure or the pressure within a container due to the compression ofgases. When a system is pulled under avacuum, the air pressure is evacuated until no air orgaseous volumes remain. This reduces the amount of pressure a compound needs to overcome toevaporate. When pressure is reduced, it, in turn, lowers the threshold of vapor pressureneeded for a compound toevaporate, escaping its liquified state into agaseous state.
This is beneficial when distilling botanical compounds because heating abotanical extractnear the atmosphericBoiling points of its target compounds for prolonged periods can induce degradation in some instances. By applyingvacuum to ourdistillation procedure, we can reduce resistance toevaporation and the amount of heat needed toevaporatethe target compound from abotanical extractaccelerating the process.Vacuum can be achieved to a varying degree depending on the strength of thevacuum pump and the design of thedistillation apparatus with nonrestrictive flow being ideal.
Vacuum is measured with any set of force per area units. In the customary US system,vacuum is measured in inches of mercury, abbreviated Hg, the atomic symbol of mercury. This measurement is determined by using a mercury barometer by measuring the distance traveled by a pool of mercury up a tube when pulled by vacuum, with the maximum level ofvacuum measured as 29.9 inches of mercury.
Within the context ofdistillation through avacuum pump, the entiredistillation apparatus is pulled undervacuum, reducing the ambient pressure of the system, which in turn reduces resistance toevaporation and the amount ofvapor pressure needed to achieve theevaporation. The continuous operation of thevacuum pump through thedistillation process also assists in pulling the ensuingvapors from theevaporator to the condenser portion of thedistillation apparatus.
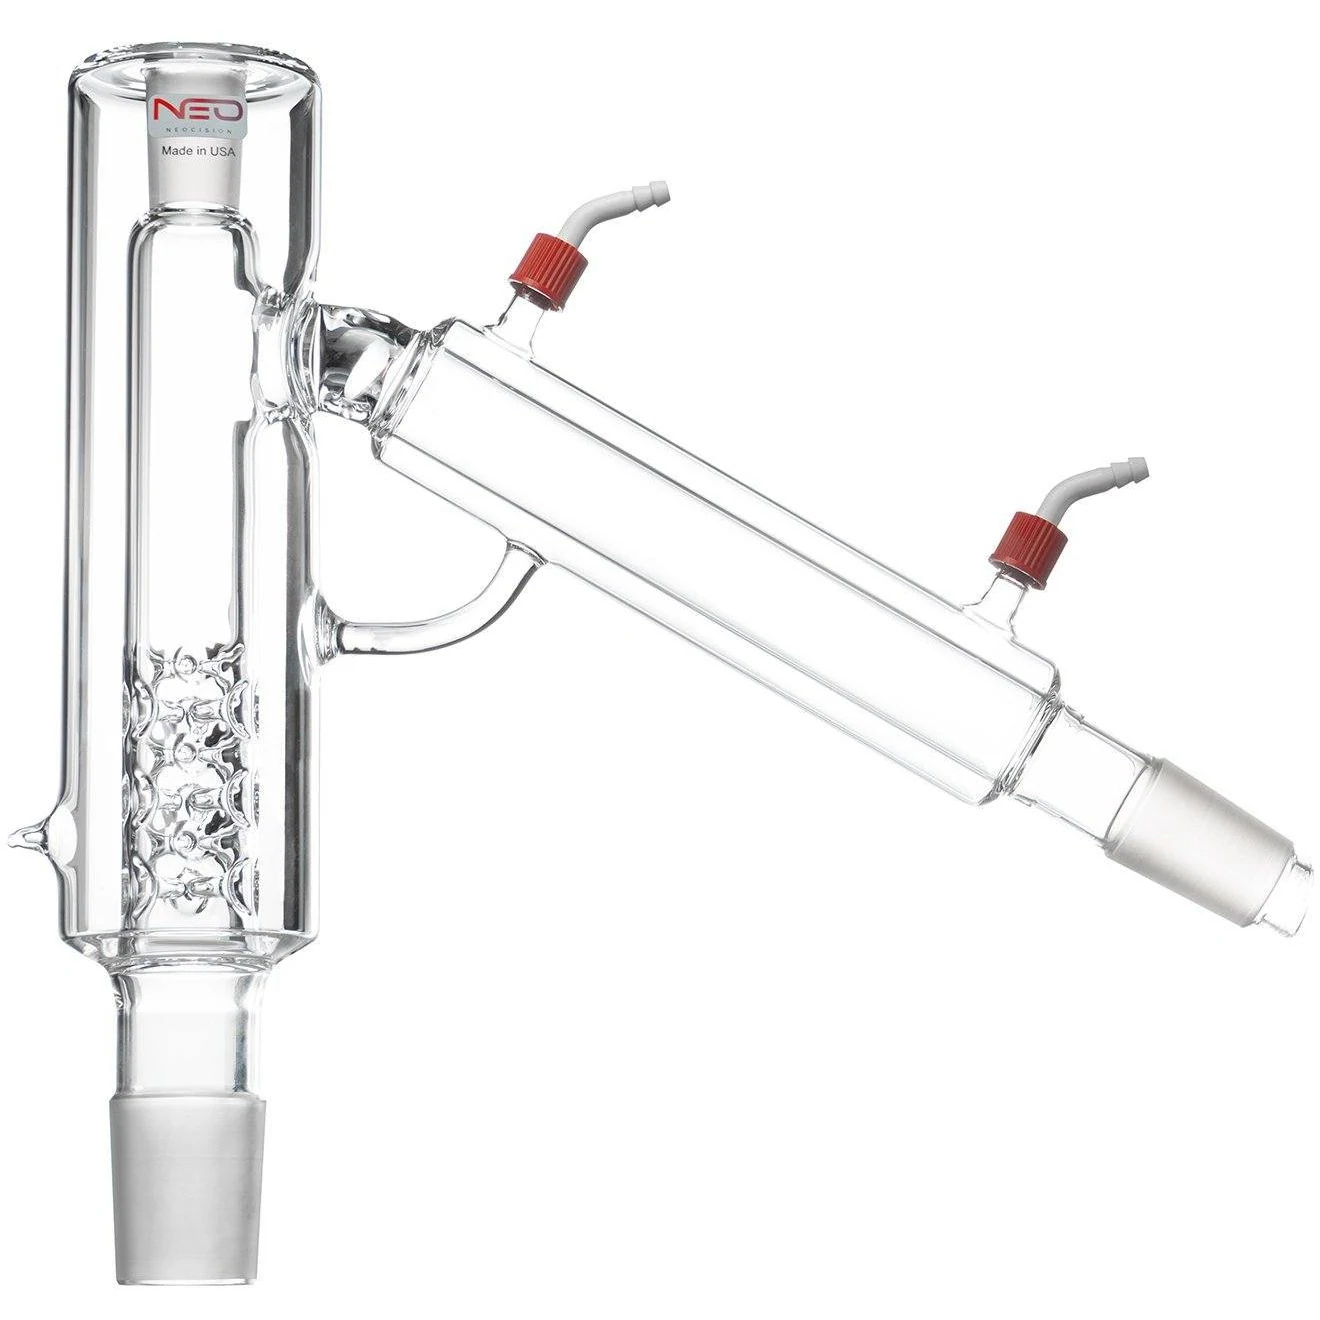
P10V6 NeocisionDistillation Head By BVV
Condensation is the other half of thedistillation process. Condensation is defined as the conversion of avapor orgas to a liquid. Condensation is initiated by the formation of molecular clusters ofvapor within agaseous volume. Condensation occurs when avapor is cooled or compressed past itssaturation limit within a given volume.
Molecules within avapor are typically far apart from one another. As morevapor collects within a given volume of air, it becomessaturated withvapor. This increase of density or closeness ofvapor molecules increases the tendency of the molecular clustering ofvapor, resulting in condensation.
Thesaturation limit of agaseous volume is the maximum amount ofvapor a given volume of air can contain. Thesaturation of agaseous volume can be achieved via eitherevaporation or cooling.Evaporation achievessaturation of agaseous volume by increasing the level ofvapor in a given volume. As morevapor collects within a given volume, that volume reaches its maximumvaporcapacity and is considered fullysaturated withvapor.
Conversely, cooling can be utilized to reduce thesaturation capacity of agaseous volume and reach itssaturation limit. Similar to thesaturation of asolvent, agaseous volume has a reducedsaturation capacity at colder temperatures. Since agaseous volume has lesssaturation capacity at lower temperatures cooling can be utilized to reach agaseousvolume'ssaturation limit and induce the molecular clustering ofvapor.
When a given volume of air issaturated withvapor, thevapor density is at its maximum. With thevapor packed so densely, condensation is likely to be initiated by the formation of molecularvapor clusters of a compound within itsgaseous volume or at the contact ofvapor with a colder liquid or a solid surface. When asaturated gaseous volume is cooled past itssaturation point, it tends to slow the movement ofvapor and initiate the clustering of molecularvapor within thegaseous volume or on a surface at a lower temperature than itself, resulting in thevapor being condensed back into a liquid phase.
Valved Vacuum Adapter 2.0 By BVV
Based on the principles of psychrometry, thevapor’s condensation point present in a sample of air can be defined as the temperature at which thevapor changes into a liquid. The temperature value that allows this process is referred to as the dew point. The dew point ofvapor is the temperature at whichvapor will condense out of thegaseous phase into the liquid phase. In the context ofdistillation,vapor does not need to be chilled excessively to initiate condensation. Generally, any temperature at least 40C/104F degrees colder than the temperature at which the compoundevaporated is sufficient In recondensing avapor back into a liquid.
During the condensation ofvaporinto a liquid, as thevapor cools thegaseous volume reaches its maximumvapor capacity or condensation point and condenses into a liquid. In doing so, the latent heat is released by thevapor and transferred to the surrounding environment. In the context ofdistillation, the heat is absorbed by the transfer fluid of the condenser. Adistillation condenser is typically a jacketed portion of thedistillation apparatus circulated with chilled fluid creating a lower temperature portion of the unit wherevapors can be condensed back into liquid form.
The condenser cools the ensuingdistillation vapors down, slowing their movement, initiating the molecular clustering ofvapor, and providing surface area forvapors to condense on. Asvacuum pulls thedistillation vapors through the condenser, the recirculated fluid absorbs the heat from thevapor, thus reducing itssaturation capacity beyond the condensation point resulting in the condensation ofvapor into a liquid. The resulting liquid formed from thedistillation process is referred to asdistillate. The condenseddistillate flows down from the condenser into a collection apparatus. Once the desired target compounds have been distilled, the result should be a highly purified extract containing upwards of 90% of the target compound. This highly pure extract can be used to make a variety of different products or be used as input for furtherisolation of target compounds.
SHORT PATH DISTILLATION
5L NeocisionShort Path DistillationTurnkey Kit By BVV
Short Path Distillationis a relatively low-costdistillation method that allows the user to collect separate fractions ofdistillate resulting in a versatile high purity botanical concentrate. Short path distillationutilizes a short distance toevaporate and recondense target compounds from abotanical extract. It is performed by applying heat andvacuum to a botanical extracttoevaporate target compounds from the startingsolution. The resultingvaporis then pulled by thevacuumpump's suction from the boiling flask, through thedistillationhead into the condenser wherevapors are re-condensed back into a liquid form resulting in a highly concentrateddistillate.
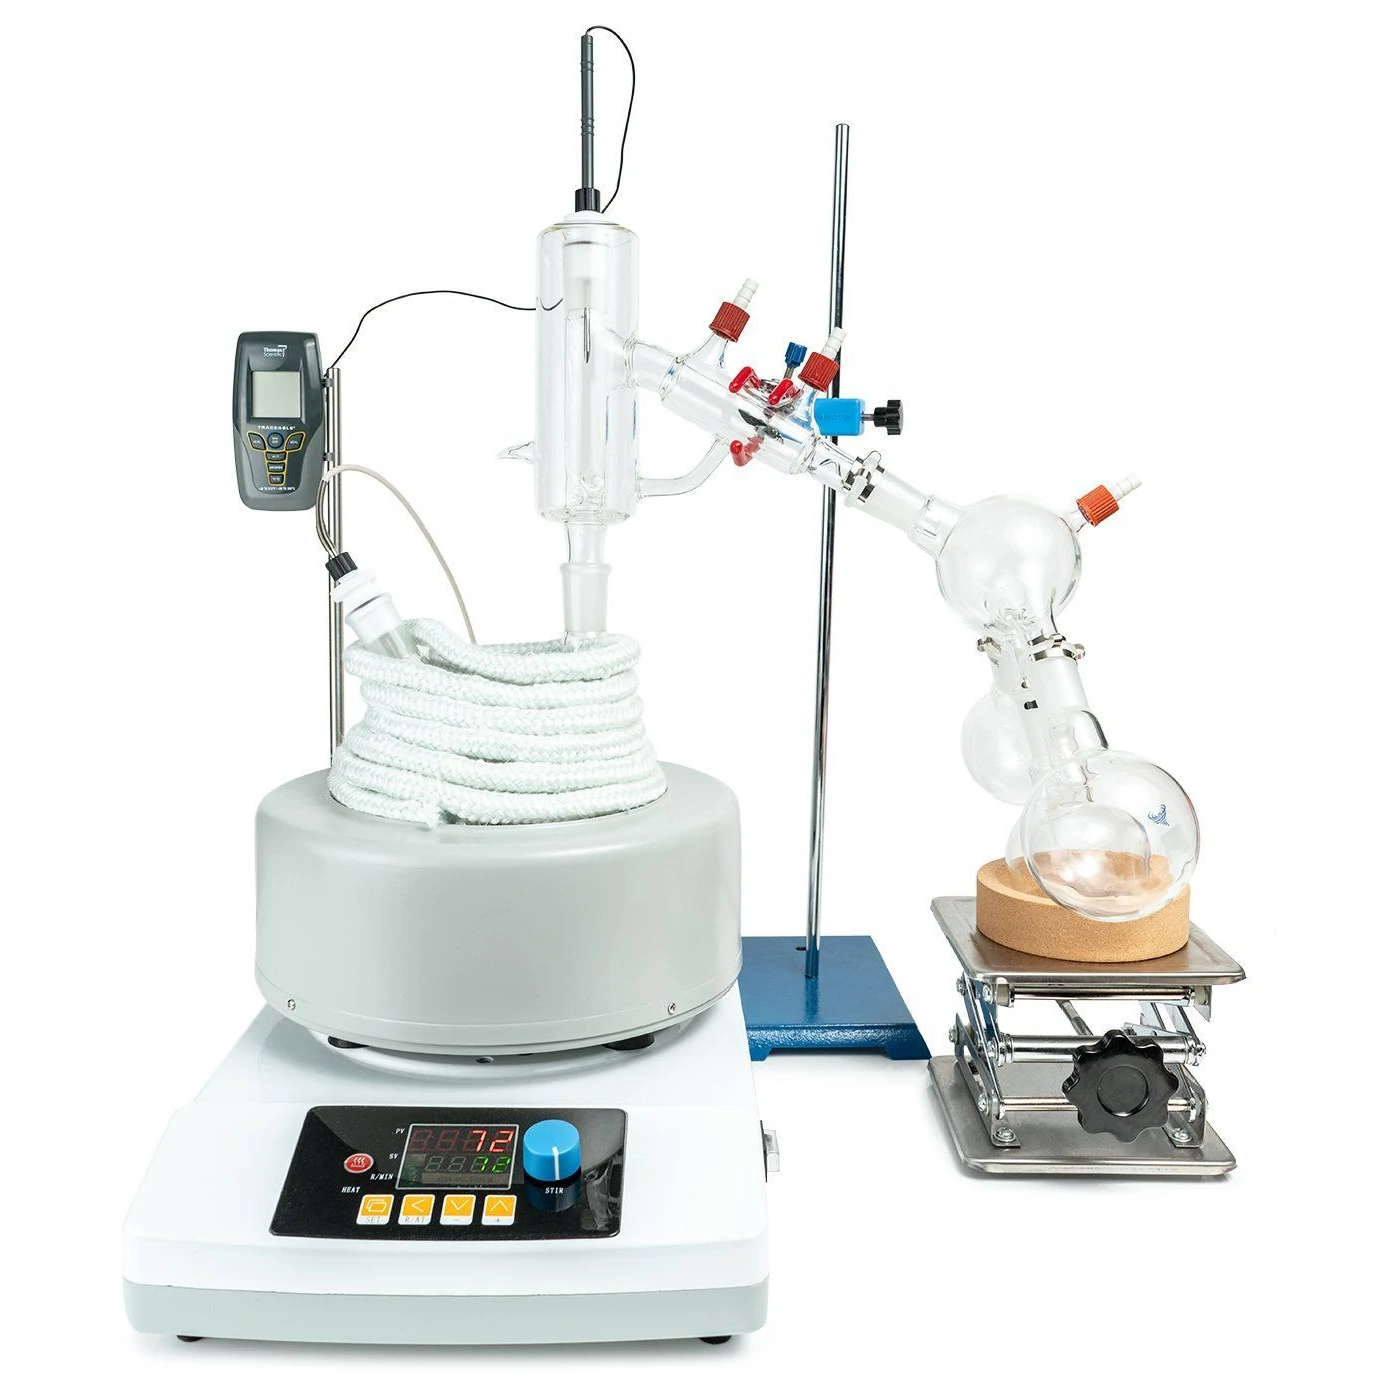
A typical short path distillation system consists of the following: a boiling flask andheated mantlewhere the input material is heated to the point ofevaporation, adistillation headwherevapor is refluxed and pulled through, a condenser chilled by acirculatorwhere the ensuingvapor is recondensed back into liquid form, a receiving flask where the distilled compounds are collected, acold trapto ensure allgaseous compounds are recondensed before reaching the final component a vacuum pump, which is typically monitored by avacuum gauge.
While each component of theshort path distillation system is essential to perform the process, thevacuum pump is most influential in the overall efficiency of theshort path distillation process. Thevacuum is used to pull the ensuingvapors through theshort path distillation system and decrease ambient pressure within theshort pathdistillation apparatus. When the system is pulled undervacuum, the pressure within the system is reduced, which lowers the amount of ambient pressure a compound must overcome in order toevaporate. By pulling the system undervacuum, resistance toevaporation is drastically reduced, increasing the efficiency of theevaporative process.
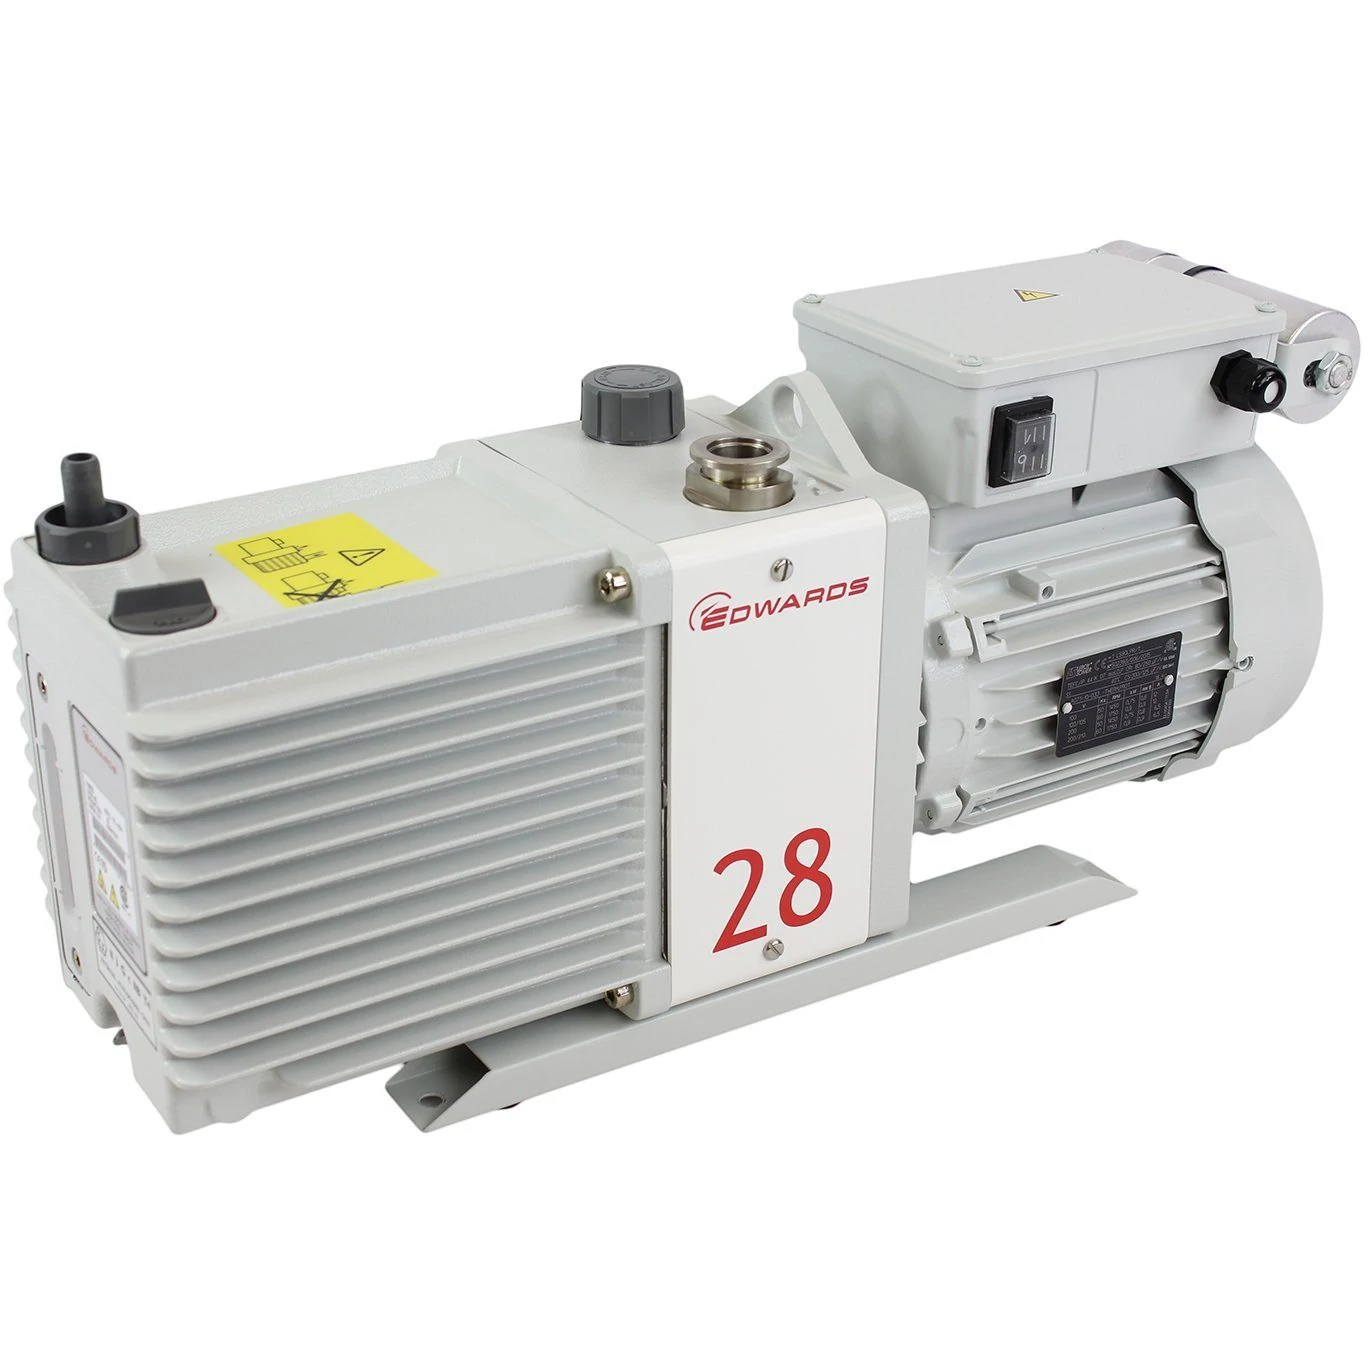
21CFM Dual-Stage High CapacityVacuum Pump By Edwards
The level ofvacuum depth ashort path distillation apparatus can achieve is attributed to the performance and strength of thevacuum pump, along with the proper assembly & configuration of theshort pathdistillation system. The configuration of theshort path distillation system should provide minimal restriction tovapor flow, high CFM from thevacuumpump, and extremely low cold trap temperatures. Beyond configuration, if thevacuum pump oil should be fresh and adequately warmed up and the system has been properly assembled with novacuum leaks,Short path distillation setups can achieve extremely lowvacuum levels duringdistillation.
The performance of thevacuum pump duringdistillationcan also be attributed to the effectiveness of the cold trap. Before thevacuum pump, the cold trap is the final component utilized to recondense anyvapors that were not condensed by the initial condenser. A cold trap is typically filled with a dry ice/ethanol slurry or achilled immersion probeto create an extremely low-temperature zone that is meant to recondense anyvapors that make it past the initial condenser. This step ensures novapors make their way into thevacuum pump, potentially contaminating thevacuum pump oil and reducing the pump's overall level of performance. Ideally, the cold trap should be chilled as cold as possible, with an ideal temperature being around -80C/-112F
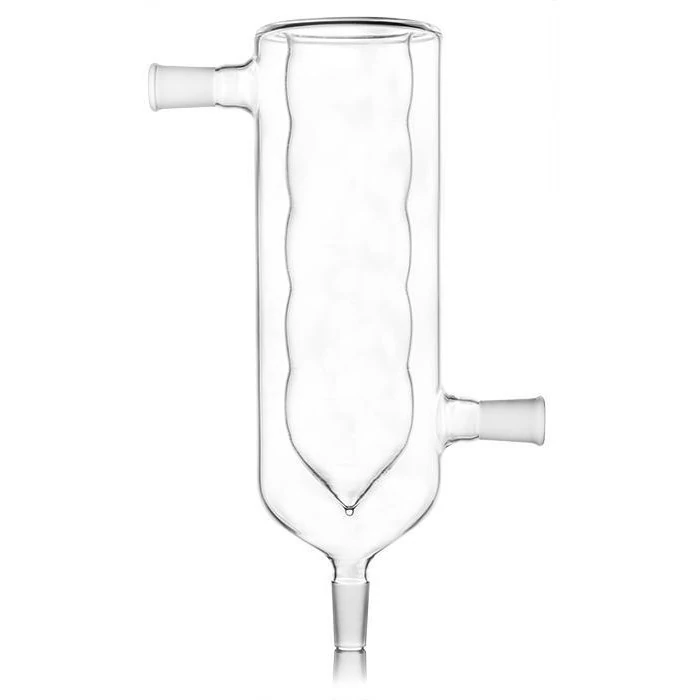
Ribbed Glass Cold Trap By BVV
While the performance of the cold trap, the seal of theshort path distillation apparatus, and the quality and temperature of thevacuum pump oil play a role in the achievablevacuum depth, these factors are limited by the strength of the short path distillation vacuum pump. Typically arotary vanevacuum pumppulls theshort pathdistillationapparatus undervacuum due to its ability to achieve deepvacuum levels. To ensure a rotary vanevacuum pump is operating at peak efficiency and qualitydistillation is achieved it is best to utilize freshvacuum pump oilfor everydistillation run as contaminatedvacuum pump oil can prevent avacuum pump from reaching the deepvacuum levels that are ideal forshort pathdistillation.
PremiumVacuum Pump Oil By BVV
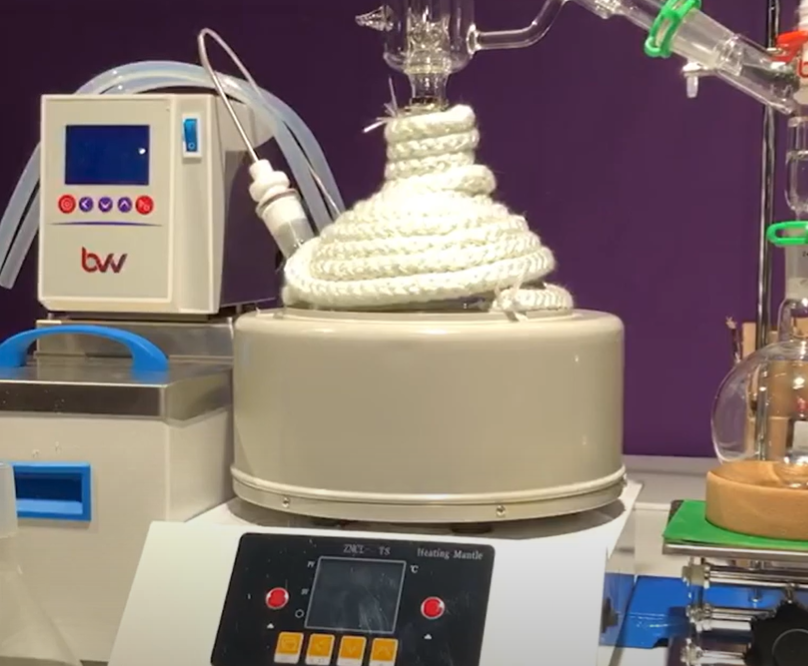
Thevacuum pump works with the heated mantle to perform theevaporative process duringdistillation. Whilevacuum drastically increases the efficiency of theevaporative process, energy in the form of heat is necessary in order to trigger theevaporative process ofdistillation. Duringshort path distillation,evaporative energy is produced by the heated mantle. The heated mantle is an apparatus designed to house and efficiently apply heat to the boiling flask. The heated mantle can be controlled to deliver precise heat to the boiling flask by monitoring the internal temperature of thesolution through the use of a thermal probe that is inserted into the boiling flask and submerged into thesolution. The thermal probe allows for precise heat application to the startingsolution allowing for controlledevaporation of target compounds.
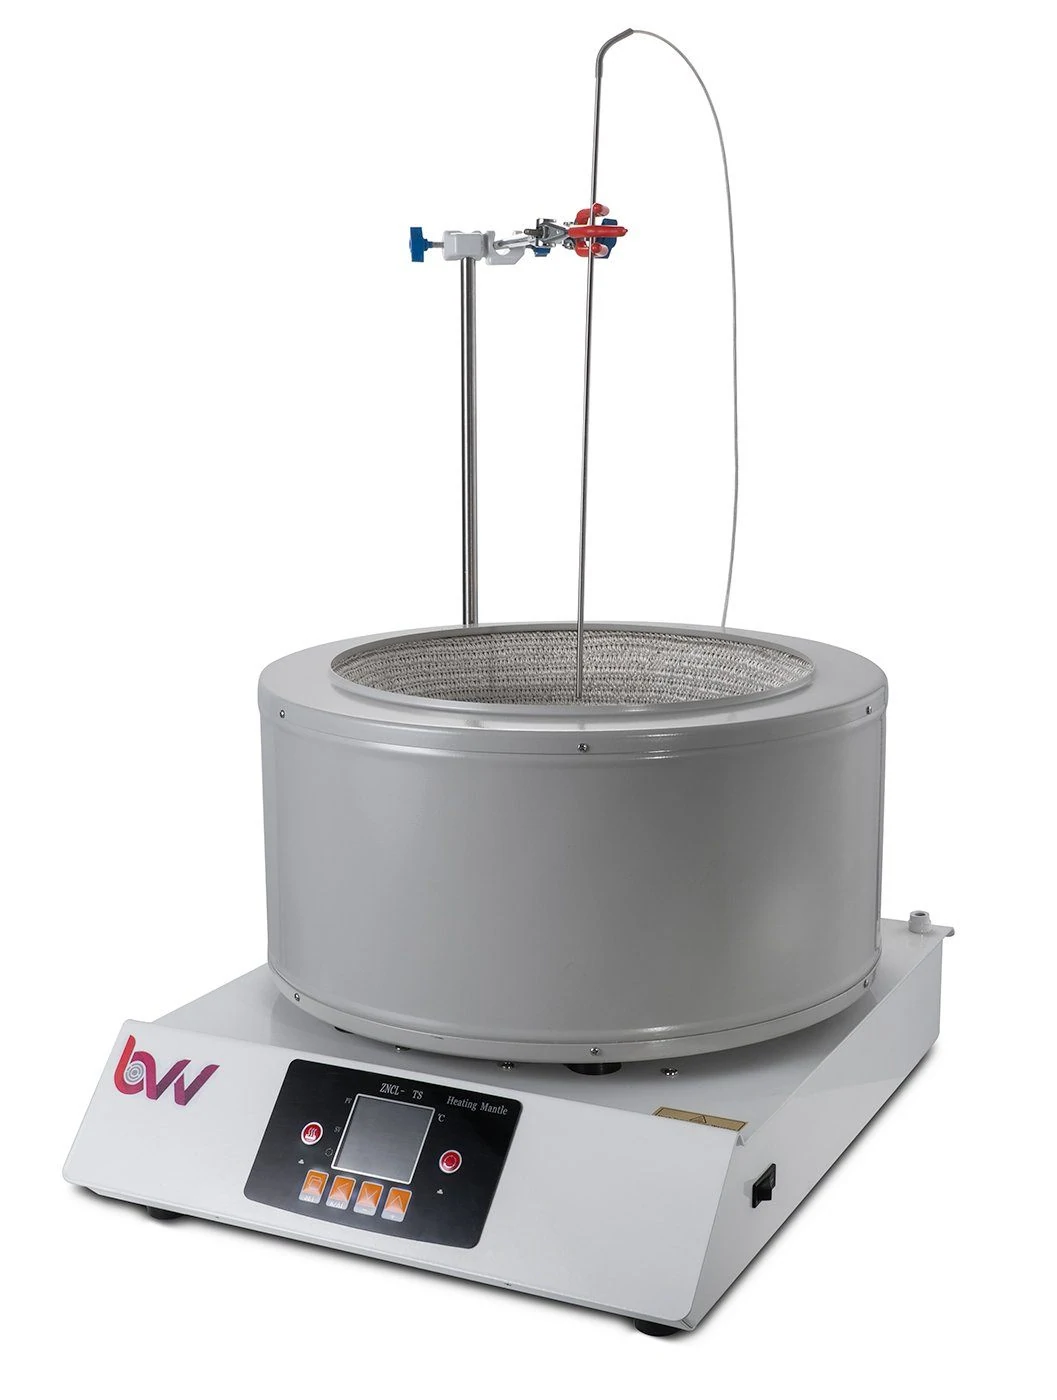
20L Digital Heating Mantel By BVV
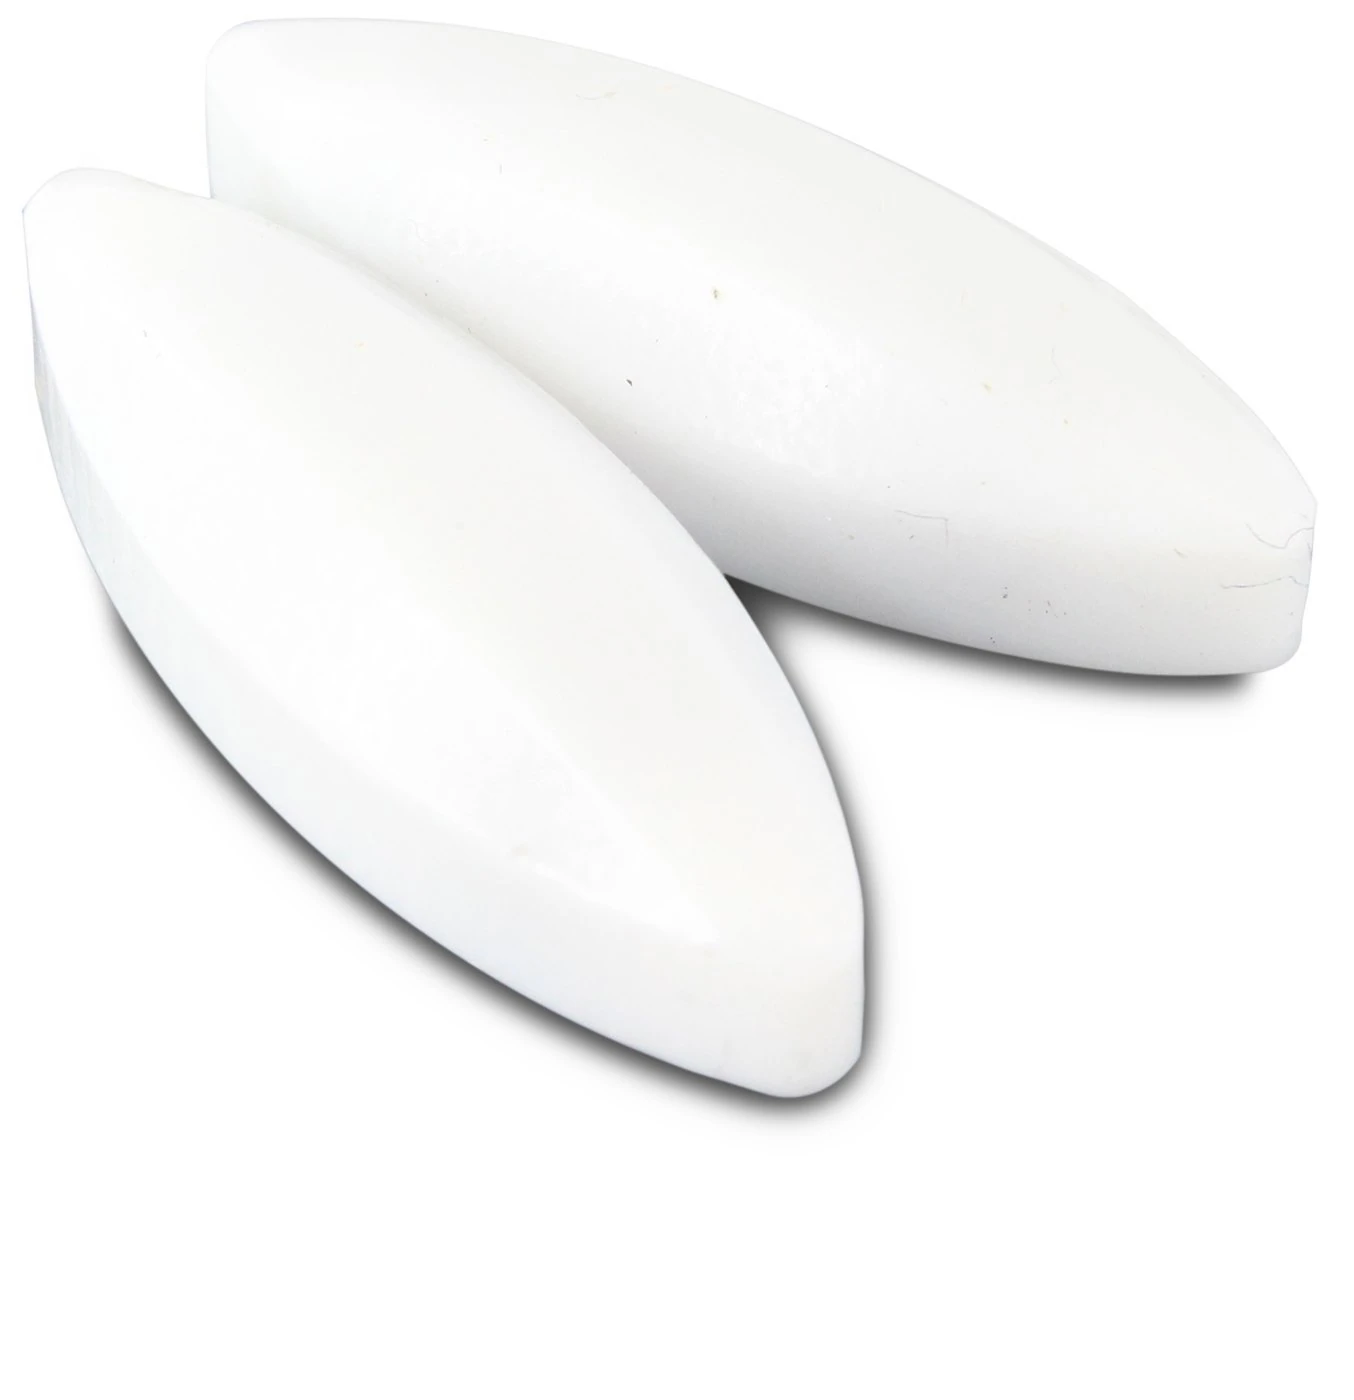
The heated mantle commonly performs both the essential application of heat to thedistillation process and provides agitation of thesolution through magnetic stirring. Heated mantle typically comes equipped with a magnetic stirring function that allows for control over the heating and agitation of thesolution within the boiling flask. This is accomplished through the use of amagnetic stir barthat is submerged into thesolution during the loading of the botanical extractinto the boiling flask. The heated mantle controls the rate at which the magnetic stir bar rotates, which results in how much thesolution is agitated. Agitation of thesolution increases heat transfer,helpsvapor bubble formations escape from thesolution more easily and attributes tovapor momentum to some extent. As a compound isevaporated from asolution, bubble formations deeper within the liquid, typically require more energy to overcome the higher fluid pressure. When agitation is applied to thesolution, the rate at whichvapor bubbles escape thesolution is accelerated, and the rate of boiling is increased.
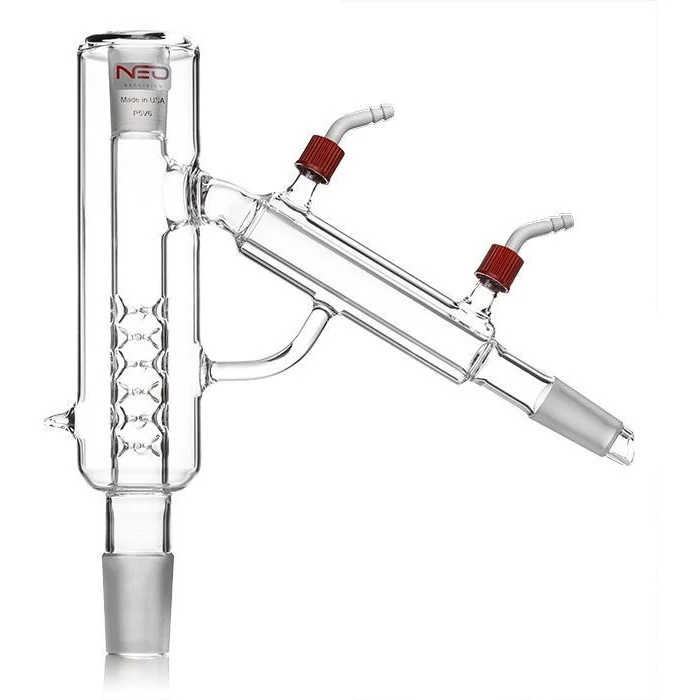
P5V6 NeocisionDistillation Head By BVV
As thesolution boils, the ensuingvapors are pulled via the suction of thevacuum pump out of the boiling flask into thedistillation head. Thedistillation head typically contains a complex pathway of glass vigreuxs that act as a "theoretical plate" or section of friction where heavier compounds with lessevaporative energy can attach to, recondense, and fall back into the boiling flask, which is referred to as reflux. The refluxing of heavier compounds with lessevaporative energy results in a purervapor stream of lighter compounds, with great enoughevaporative energy to pass through thedistillation head into the condenser.
Traceable Kangaroo Digital Thermometer By Thermo Scientific
While the primary function of thedistillation head is to accomplish the separation of heavier and lightervapors to create a more purevapor stream and resultingdistillate flow, thedistillationhead also serves as a point wherevapor temperature can be measured through the use of athermal probe.Distillation heads commonly contain a port where a thermal probe can be inserted, allowing for the invaluable measurement of thedistillation headvapor temperature.Distillation headvapor temperature Is an incredibly valuable tool in measuring theevaporative process. Observation of a drop in headvapor temperature or an observed decreased boiling rate is a great indicator thatevaporation at the set temperature has decreased and mantle temperature can be increased to continue theevaporative process.
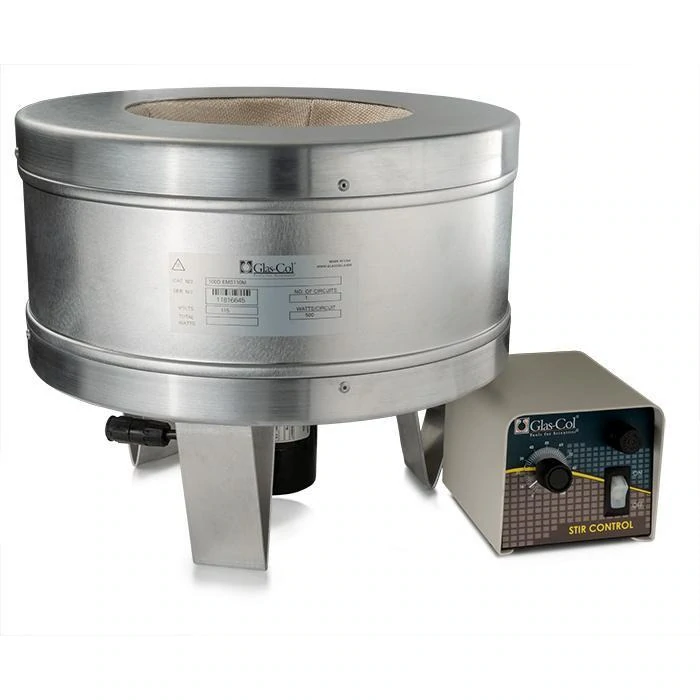
Stirmantle By Glas-Col
During theevaporative process, as mantle temperature is increased, it subsequently increases headvapor temperature. Over time as thesolution boils at the set mantle temperature, the compounds with correspondingvapor pressureevaporate from thesolution resulting in a reduction ofvapor being created over time, which can be observed visually as a decreased rate of boiling and an overall drop in headvapor temperature. When a 0.5C decrease in headvapor temperature is observed, it can be used as an indicator that the rate ofevaporation of compounds in that temperature range has decreased, and mantle temperature can be increased to continue theevaporative process and ensure a timelydistillation. By incrementally increasing mantle temperature based on dips indistillation headvapor temperature, the operator can achieve the most efficientdistillation possible.
Conversely, an experienced operator can establish timed intervals to increase temperatures or increase the mantle temperature as it warms by defined increments.Short path distillation can be performed successfully with various methodologies that are commonly centered around increasing mantle temperature incrementally to separate compounds via selectiveevaporation, condensation, and collection.
P10V6 Neocision Distillation Head - USA Made By BVV
Duringdistillation, the ensuingvapors that make their way through thedistillation head arrive at the condenser portion of the short path distillation apparatus. This jacketed glass portion of theshort path distillation system allows for heated or chilled thermal fluid to be recirculated through the jacket utilizing aheated circulator. This lower temperature portion of the short path distillation apparatus is utilized to recondense thevapors that make it through thedistillation head so the resultingdistillate can be collected into the receiving flask.
Contrary to popular belief, the condenser does not need to be extremely cold to effectively condensevapors; typically a 20C/68F colder condenser temperature than thevapor temperature is sufficient in condensingdistillate vapors. While cold condenser temperatures can be ideal when distilling volatile compounds liketerpenes, too cold of a condenser can increase theviscosity of thedistillate stream, which can result in the clogging of the condenser. Conversely, too hot of a condenser temperature can result invapor flowing through the condenser without being condensed.
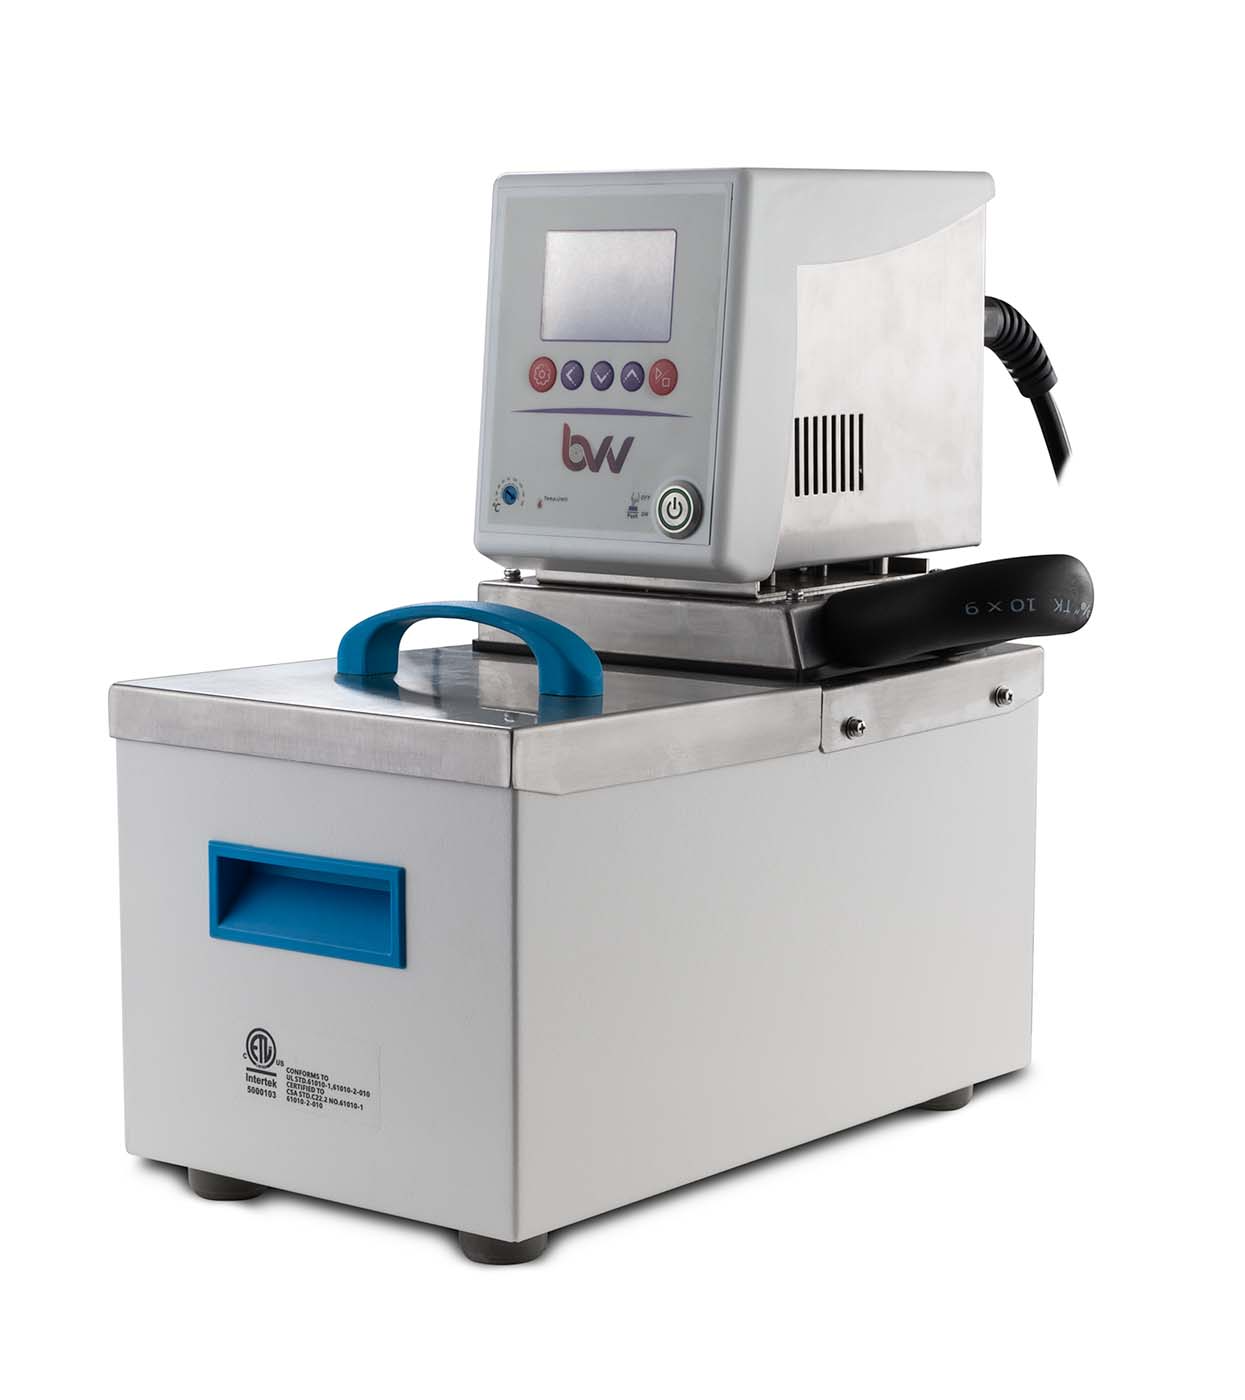
5 Liter Neocision Heated Circulator By BVV
Duringdistillation, the condenser circulator temperature can be set using a variety of different strategies, including leaving the condenser at the same temperature through the entirety of thedistillation procedure or incrementally increasing the circulator temperature to maintain a 20-degree colder temperature than thevapor temperature up to the heated circulator’s limit and lines while not exceeding a heated recirculation temperature of 130C/266F. While both methods can produce excellent results, the latter is recommended for novice operators with an ideal heated circulator temperature range between 70-90C/158-194F being sufficient for condensingvapors through the entirety of thedistillation.
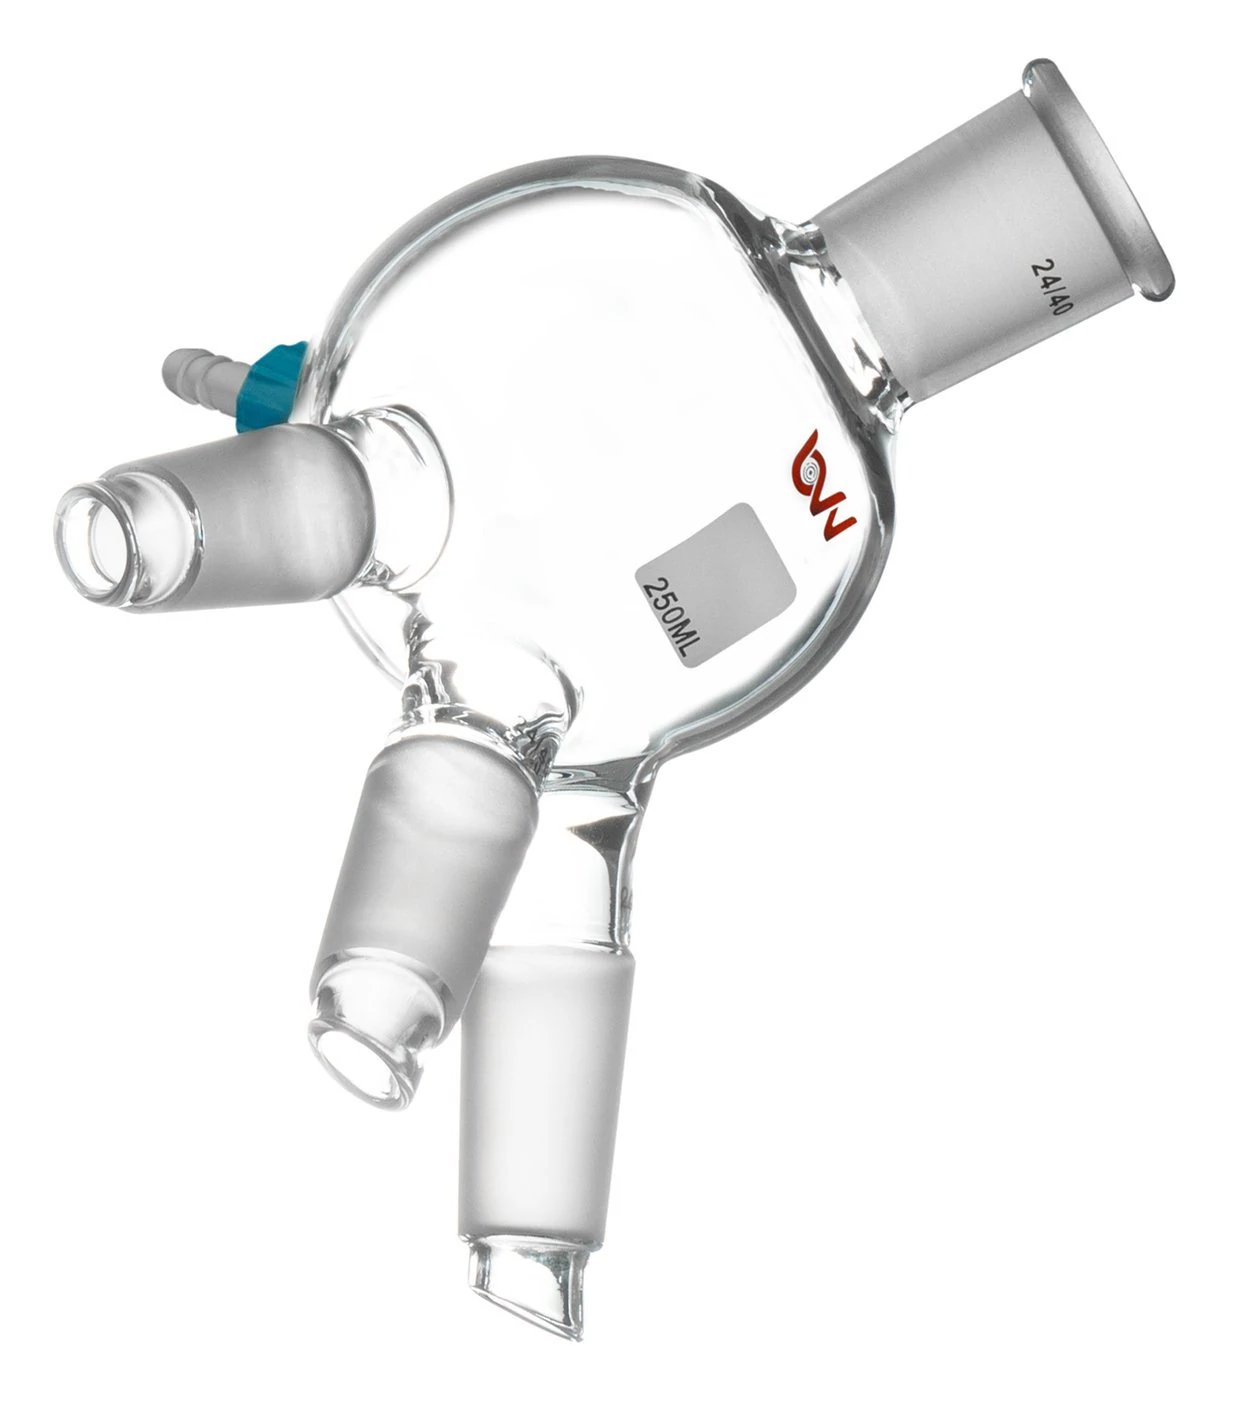
Depending on the short path distillation apparatus after the condenser, there can be a variety of different collection apparatuses utilized for capturing different fractions from the distillation process ranging from a single collection vessel with an isolation valve, a multi-position cow, a swing arm adaptor, or a discharge pump. While swing arm adaptors and multi-position cows are great for collecting multiple fractions without breaking vacuum, isolation valves and discharge pumps allow for the capturing an infinite amount of fractions during distillation with a discharge pump allowing for ultimate efficiency for high volume distillers. Regardless of the collection apparatus, the goal of all these components is to allow for the collection and separation of distillate fractions.
Outlet Pump By Beaker & Wrench
Short path distillation is commonly used as a form offractionaldistillation by incrementally increasing the heat applied by the mantle to distill and separate compounds based on their difference inboiling points and orvapor pressures. The boiling pointof a substance is the temperature at which a pure substanceevaporates, observed as boiling under normal atmospheric conditions. When dealing with a mixture of many compounds like that of abotanical extract, each compound contributes a fraction of the totalvapor pressurebased on their respectiveboiling points, known as partialvapor pressure.
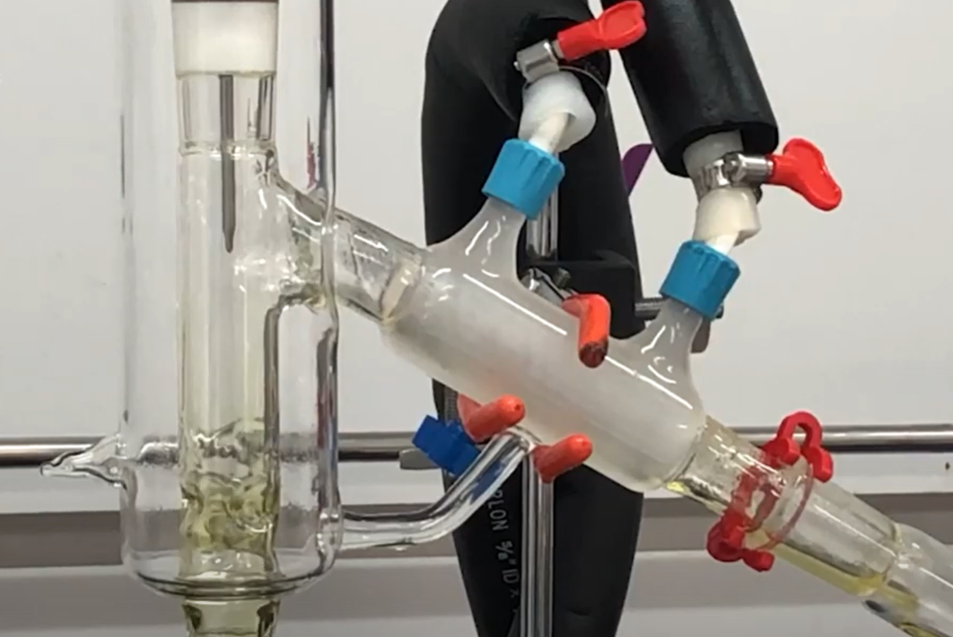
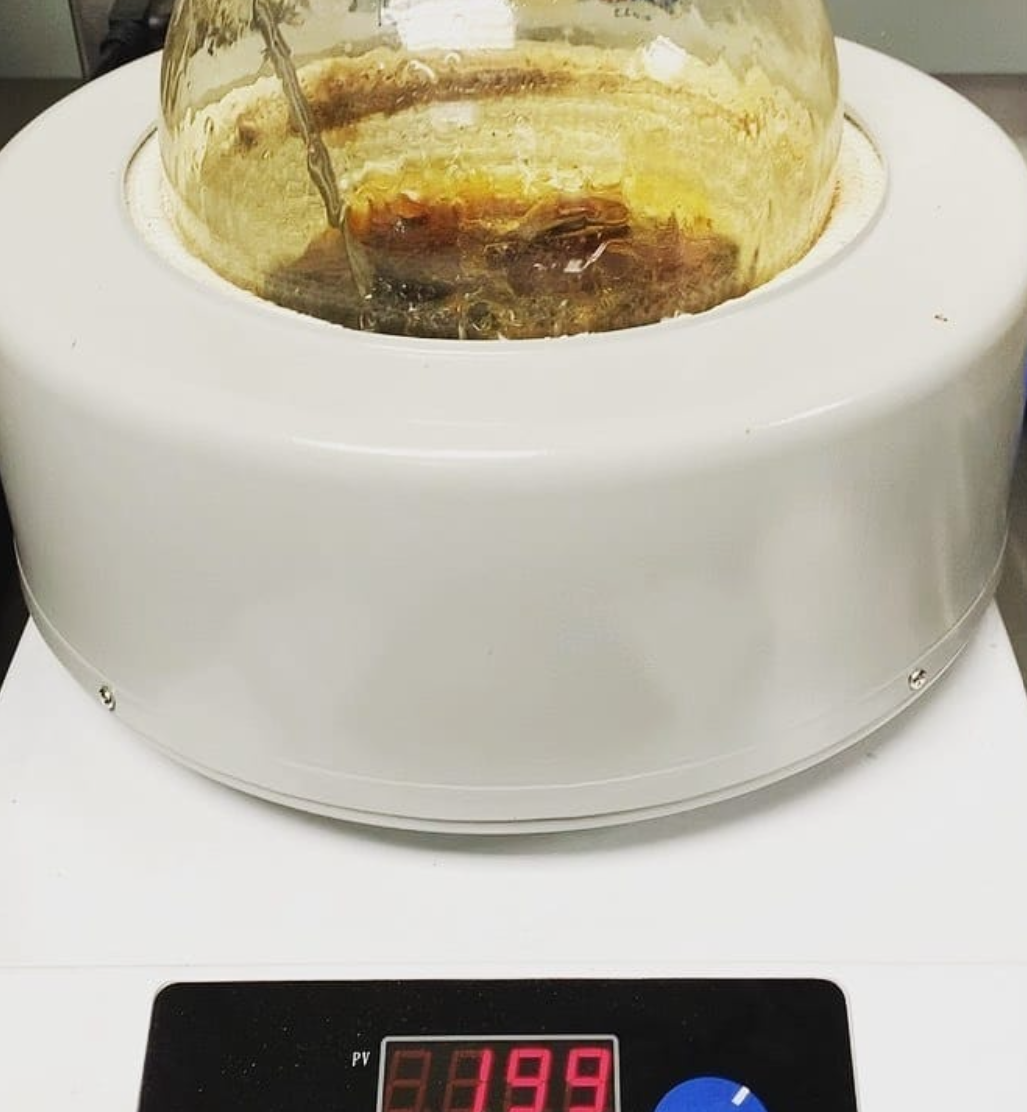
Evaporation inside a boiling flask
Typically the lower a compound'sboiling point, the higher itsvapor pressure. Once thevapor pressureof a compound exceeds the ambient pressure of the closed system containing it, that compound willevaporate from thesolution observed as boiling. As the temperature of thesolution is increased,vapor pressureis also increased, resulting in compoundsevaporating from thesolution in order based on their respectiveboiling points, starting with the most volatile compound with the lowest boiling points.
During distillation, compounds with similar boiling points tend to evaporate within the same temperature range, making it difficult to completely isolate compounds. While complete isolation of a certain compound is typically easier to achieve through crystallization or chromatography. Fractional distillation can be utilized to efficiently separate compounds with different boiling point ranges and collect highly pure fractions of compounds that evaporate within the same temperature range.
2LShort Path Distillation Kit By BVV
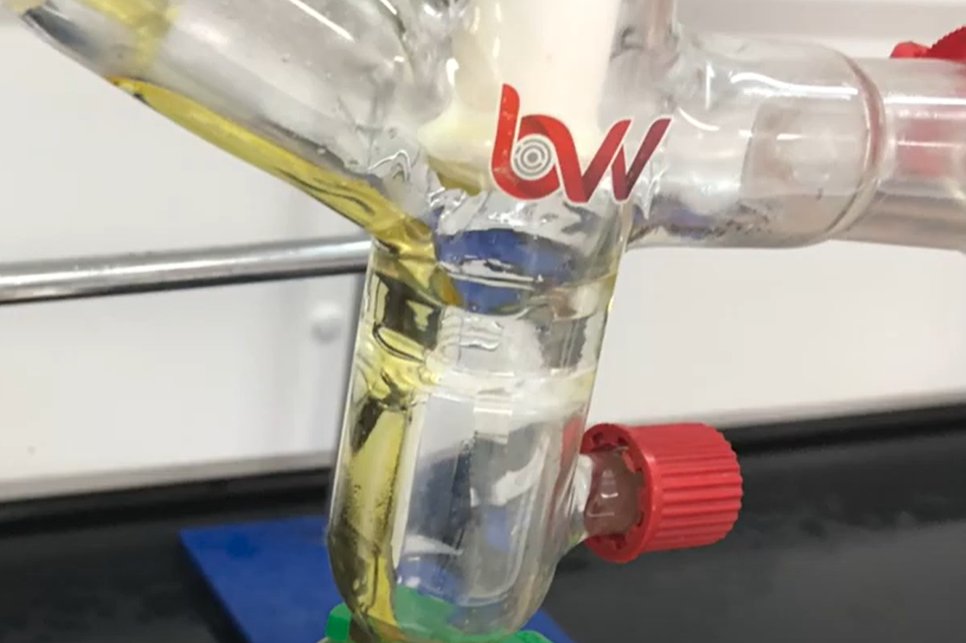
Typically, these fractions are collected into three categories: the heads fraction, the main body fraction, and the tails fraction. The heads fraction contains the compounds firstevaporated from thesolution, consisting of the compounds with a lowerboiling pointthan that of the target compounds. As the heated mantle temperature increases and the target compounds start toevaporate from thesolution, the main body fraction starts to distill. At this point of thedistillation, the collection or receiving flask is changed in order to separate the lowerboiling pointcompounds within the heads fraction from the target compounds of thedistillation to collect a higher purity fraction of the target compounds referred to as the main body.
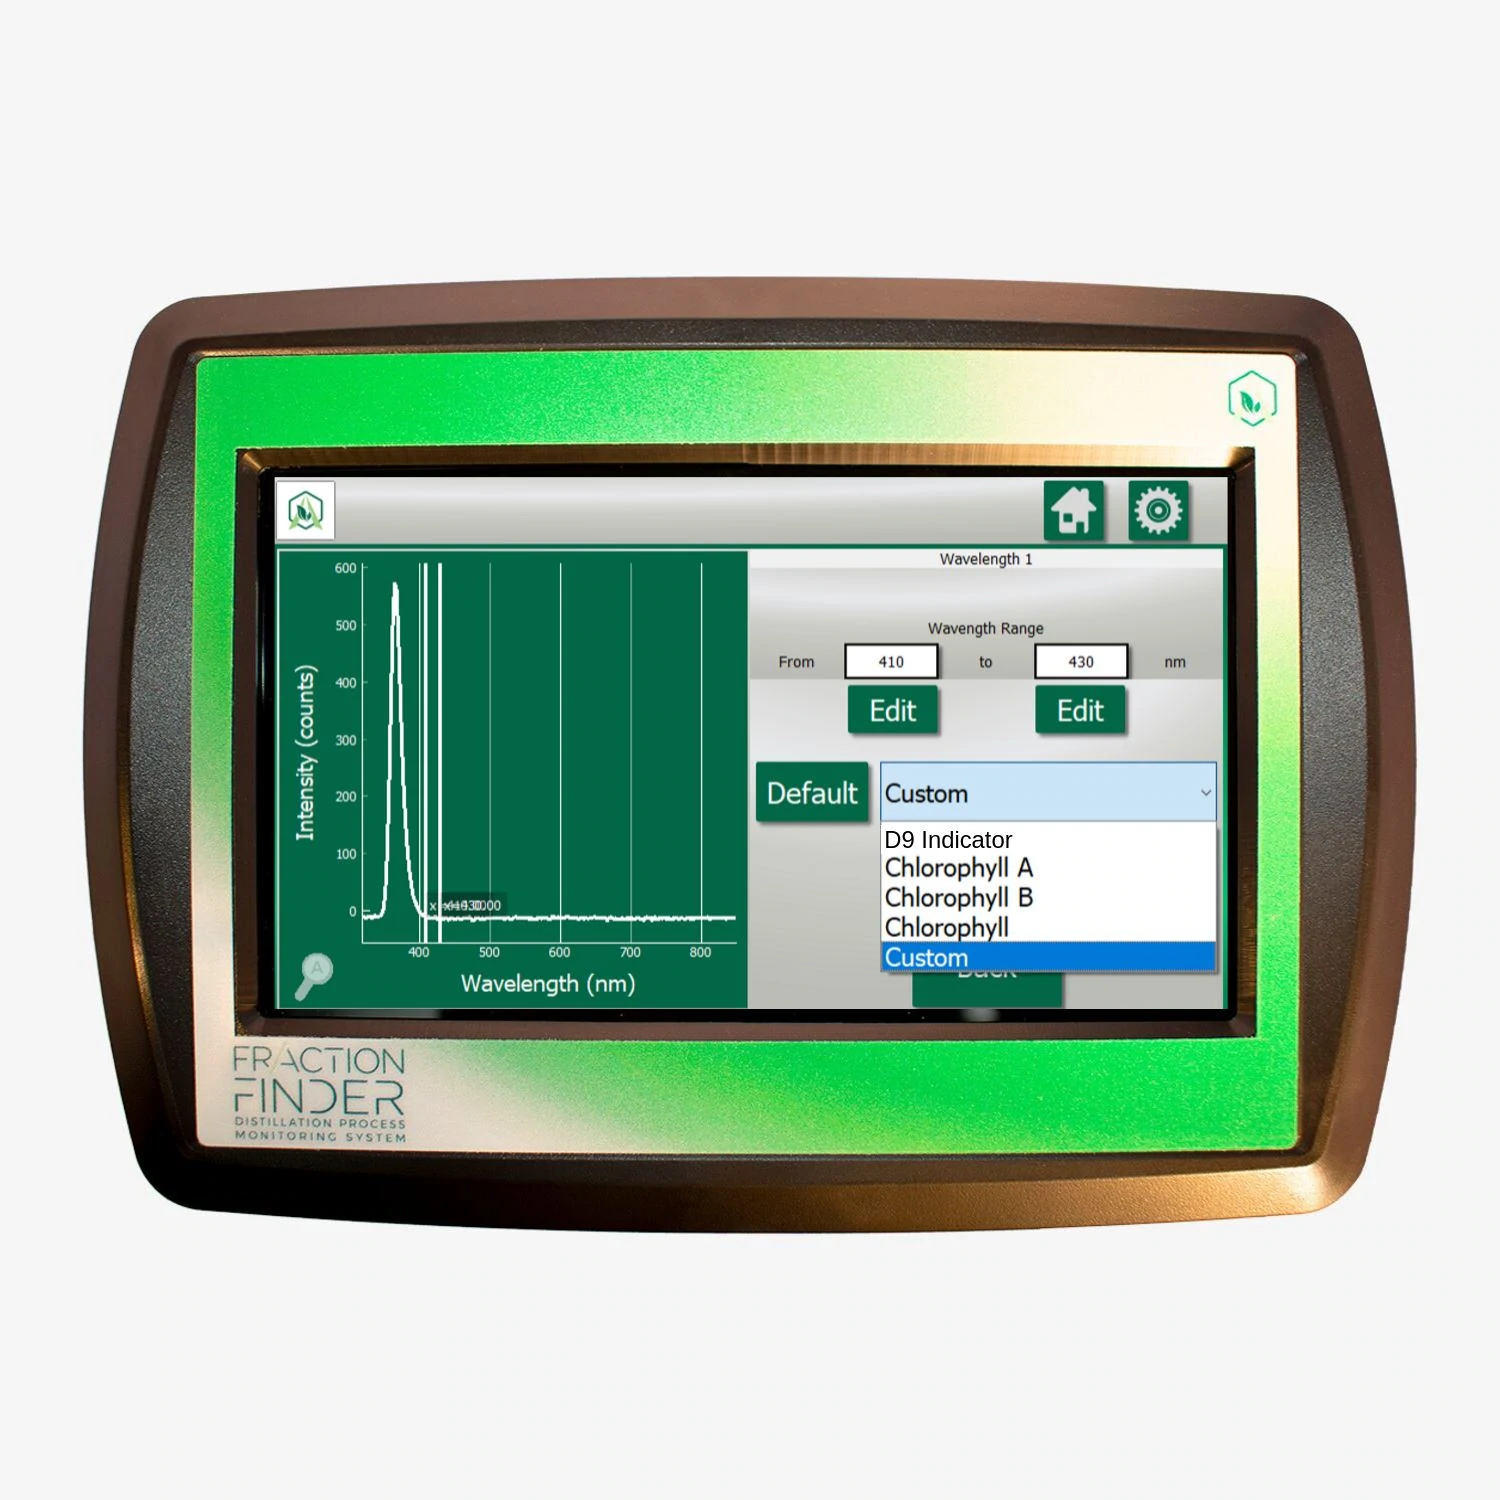
Fraction Finder By Arometrix
Duringdistillation, it can be challenging to pinpoint the appropriate time to change flasks between the heads and main body fraction. While aUV Detectorcan be utilized to pinpoint which compounds are being distilled without this type of device, it is commonly left to the observation ofdistillate flow. Typically Ifdistillate viscosity is increased or If analytical results show that the starting input predominantly consists of the target compound and an increased flow ofdistillate is observed, both can be a good indicator that the target compounds are starting to distill and the collection flask should be changed to isolate the main body from the heads fraction.
Heads Fraction
To ensure the main body fraction is of the highest purity, it is common to allow some of the main body to collect into the heads fraction before changing flasks. This ensures that the lighter volatile compounds are not collected into the main body fraction. Once the main body is assumed to be distilling before swapping the flask, it's best to allowevaporation at that temperature range to diminish, observed by a significant drop in headvapor temperature and a visual reduction of boiling to ensure the lowerboiling pointcompounds have distilled before swapping collection flasks and increasing mantle temperature toevaporate the target compounds of the main body fraction at a higher temperature range.
As thedistillation process is resumed and theevaporation of the main body is achieved. Once satisfied with the collection of the main body,distillation can be stopped leaving the tails fraction remaining in the resinous remains of the starting material, or the main body flask can be changed, and the tails fraction can beevaporated from thesolution into a separate collection vessel. While the tails fraction typically contains some of the target compounds asdistillation temperatures increase, color pigments tend to darken thedistillate flow that most distillers prefer to isolate from the main body fraction in order to keep the main body of the highest purity.
Fractional distillation is a process refined by the operator over time. While the ideal separation ofdistillate fractions can be hard to pinpoint, the process can be optimized through analytical testing, trial, and error. At the same time, the separation of fractions is dependent on the target compounds of thedistillation. As a general rule of thumb, an increased rate ofdistillate flow andviscosity are good indications thedistillation is entering the main body fraction, and a visual darkening of thedistillate flow can indicate the tails fraction is starting to distill.
Before the process of short path distillation can be initiated, the apparatus must first be appropriately staged and properly assembled. Ashort pathdistillation system is meant for operation within a fume hood or within a well-ventilated area. When staging ashort path distillation apparatus, assure the system is assembled in an area where thevacuum pump exhaust can be vented outside the facility.
When a suitable area has been identified, short pathdistillation assembly starts with positioning the heated mantle and support stands. For greater ease of use, the heated mantle of a short pathdistillation apparatus is commonly elevated to allow for more clearance when changing the collection flask to separate fractions. After staging the mantle, preassemble all lab stands, boss heads, three-finger clamps, chain clamps, scissor jacks, and make sure to have keck clips and vacuum grease within arms reach to aid in the assembly of glassware.
Laboratory Scissor Jack By BVV
Proper assembly of theshort path distillation apparatus is essential for producing the deepvacuum levels that are ideal forshort path distillation. To assure the deepest possiblevacuum is achieved, it is important that each joint of theshort pathdistillation system is appropriately assembled withvacuum grease to create a complete seal between joints. When choosing avacuum grease for application, ensure thegrease is rated for the temperatures of thedistillation. For temperatures above 204C/399FApizon High-TemperatureVacuum Greaseis highly recommended.
Vacuum grease is used to create a perfect seal between the separate components of theshort pathdistillation apparatus. To properlygrease each joint, place a small amount ofvacuum grease on the upper half of the male joint and insert it into the female joint. Once in place, rotate the male joint until an even coating ofvacuum grease can be seen across the entire circumference of the upper half of the male joint. When a complete seal ofvacuumgrease has been made, secure the joint with an appropriately sizedkeck clipand support the component with the appropriate lab stand support or scissor jack. Keep in mind thatvacuum grease should only be applied to the upper half of the male joint to ensurevacuum grease does not come in contact with the lower half of the joint, potentially contaminating the inside of the system.
High-TemperatureVacuum Grease By Apiezon
During the assembly process, it is important to ensure all glass joints are appropriately greased withvacuum grease, creating a complete seal, and each component is fully supported by the appropriate lab stands or scissor jacks. To start the process of assembly, first place the boiling flask inside the heating mantle, and install the mantle temperature probe and connect thedistillation head to the boiling flask. Once thedistillation head has been properly attached, install thedistillation head temperature probe and proceeds to attach the condenser if it isn't already integrated into thedistillation head. After the condenser has been properly installed and supported, proceed to assemble the remainder of theshort pathdistillation glassware prior to connecting the condenser to the heated circulator to ensure joints can be manipulated into place.
Triple neck Cow Receiving Flask By BVV
Continue assembly by connecting the condenser to the collection apparatus and installing thereceiving flasksmaking sure each component is connected properly and supported using the appropriate lab stand or scissor jack supports. After assembly of the collection apparatus, proceed to connect the cold trap ensuring all connections are appropriately greased, and the cold trap is properly supported using a chain clamp. Once the cold trap has been properly assembled, proceed to connect thevacuum gaugeandvacuum pump using the appropriatevacuum-rated lines and plumb thevacuum pump exhaust outside the facility.
Now that the short path distillation apparatus has been completely assembled before proceeding tovacuum test the system, prior to doing so, double-check to ensure allglassware is appropriately supported by the lab stands and all components are properly connected. Proceed tovacuum test the system by first ensuring thevacuum pump is appropriately filled with freshvacuum pump oil and closing thevacuum valve to the system before turning on thevacuum pump in open gas ballastmode and allowing it to warm up for 30 minutes to an hour. Once thevacuum pump oil has been appropriately warmed up, proceed to open thevacuum valve allowing the entire system to be pulled undervacuum.
Bullseye PrecisionVacuum Gauge By DigiVac
At this point, monitor thevacuum gauge.Vacuum gauges provide invaluable monitoring ofvacuum pressure duringdistillation, typicallyvacuum gauges are utilized after the cold trap, with an additionalvacuum gauge before the cold trap providing data on the effectiveness of the cold trap. While handheld gauges provide a cost-effective monitoringsolution, precisionvacuum gauges are ideal for professional distillers allowing for precise monitoring ofvacuum pressure.
During the initialvacuum pull, if deepvacuum levels are not attained, proceed to identify where thevacuum may be leaking by spraying each glass joint with soapy water, turning off thevacuum pump, and bleeding pressure before disassembling and reassembling connections that are not maintaining a seal. Proceed tovacuum test and disassemble and reassemble the system as needed until there are novacuum leaks andvacuum levels can be maintained.
Once assembly is complete, and theshort pathdistillation apparatus has been proven to maintainvacuum levels, it's best to test the achievable amount ofvacuum depth by allowing the system to be pulled undervacuum in closed ballast mode. Given the system is properly assembled, In closed ballast mode, a rotary vanevacuum pump should be able to achievevacuumdepths below 25microns with even lower depths being achievable prior to loading the system with thebotanical extract.
2L PremiumShort PathDistillation Kit By BVV
After thevacuum testing is complete, the last task is to connect the heated circulator to the condenser port and fill the heated circulator with the appropriate heat transfer fluid. If operating the heated circulator in temperature ranges below 90C/194F, a distilled water or water-glycol mixture is sufficient. If operating a heated circulator above 90C/194F, a high-temperatureheat transfer fluidlike silicone oil is recommended. When connecting the heated circulator to the condenser, ensure the heat transfer fluid inlet is flowing from the top of the condenser to the bottom, and all clamps are tightened appropriately.
Theshort path distillation system is now ready for operation. To prepare forshort path distillation, start by bringing the system to temperature and allowing thevacuum pump to warm up in open ballast mode. Again ensure thevacuum pump oil is filled to the appropriate operating level and thevacuum valve to the system is closed, before proceeding to turn on thevacuum pump in open ballast mode and allowing it to warm up for 30 minutes to an hour before operation while the rest of the system is prepared.
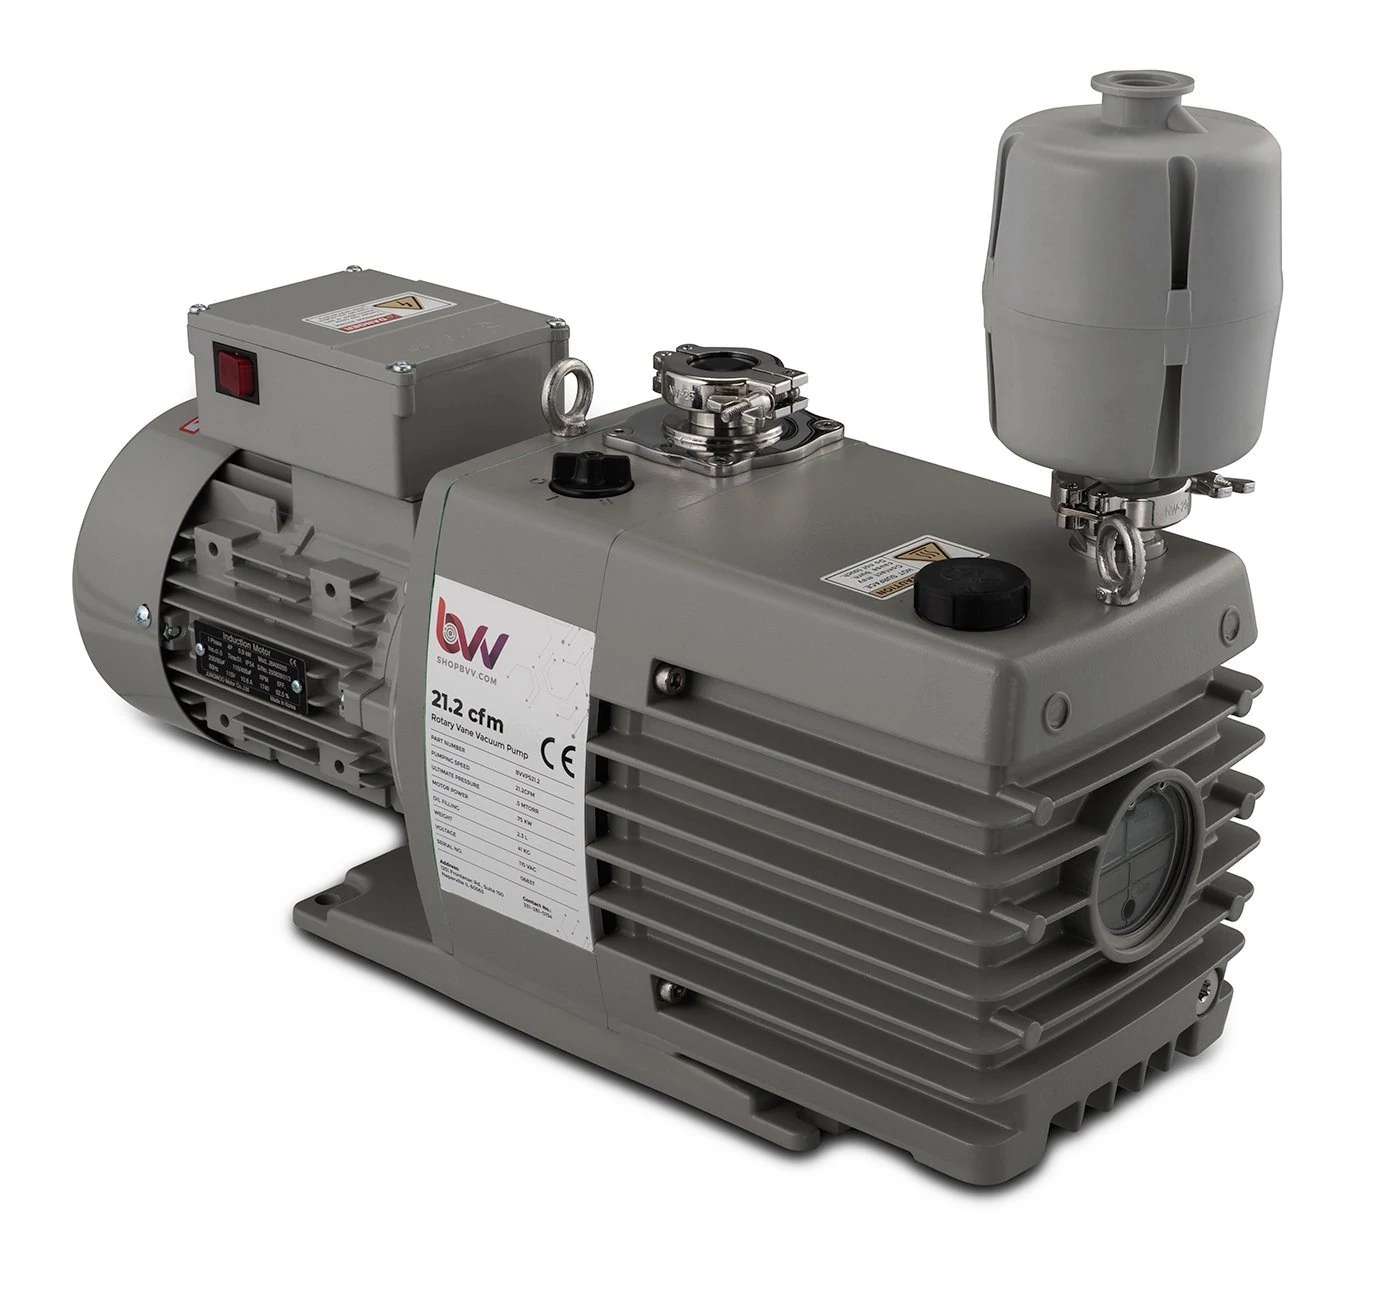
21.2CFM Corrosion Resistant Two StageVacuum Pump By BVV
While thevacuum pump is warming up in open ballast mode, proceed to bring the cold trap to operating temperature by turning on the cold trap immersion probe chiller setting it to its lowest setting, or by preparing the cold trap with dry ice andethanol slurry by filling the cold trap 1/2 way withethanol and slowly adding dry ice to avoid over the bubbling of thesolution.
As the cold trap reaches operating temperature, proceed to bring the condenser's heated circulator to operating temperature by first ensuring that the circulator bath is filled to the appropriate fill level and the fluid circulation is connected to flow into the top of the condenser and out of the bottom of the condenser. Proceed to ensure all clamps are tightened appropriately before turning on the circulator and setting it to the desired condenser temperature. For the start of the run, a temperature range between 40-50C/104F-122F is effective at condensing volatile compounds. Once volatiles have diminished a condenser temperature of 70-90C/158-194F can be used and maintained for the entirety of the run, or condenser temperature can be increased from this point to maintain a 20C/68F degree colder difference between thevapor temperature and the condenser temperature.
Digital Heating Mantle Probe By BVV
As thevacuum pump warms up and the system reaches the desired operating temperature, proceed to load thebotanical extractinto the boiling flask by removing the mantle thermal probe, cleaning the joint of anyvacuum grease, and attaching a glass funnel. Proceed to fill the boiling flask with winterized and devolatilized botanical extract. The boiling flask should be filled to a minimum of 25% of the capacity of the boiling flask, ensuring that there is enoughsolution for the thermal probe to read, and no more than 50% of the total volume of the boiling flask to ensure the there is enough headspace for thesolution to boil. Keep in mind winterized and devolatilizedbotanical extractis ideal for short pathdistillation as unwinterizedcrude oilwill make cleaning the boiling flask difficult, and non-devolitized crude may impede its ability to achieve idealvacuum levels.
Fabric Insulating Top By Glas-Col
Once thebotanical extracthas been loaded into the boiling flask, add the magnetic stir bar and reinstall the mantle temperature probe into the boiling flask ensuring the probe reaches 1/4" from the bottom of the boiling flask without touching the bottom of the flask or interfering with the rotation of the magnetic stir bar. The system is now sealed and can bevacuum tested once again to ensure a deepvacuum is reached before applying heat to the system. Proceed to switch thevacuum pump into closed gas ballastmode and slowly open thevacuum valve to pull the system undervacuum. Ideally, avacuum range below 50microns is ideal, with the deeper thevacuumdepth the better. At this time before heat is applied to the mantle it is highly recommended to apply insulation to the boiling flask utilizing an appropriate boiling flask jacket orinsulation ropeto ensure boiling flask temperatures are maintained duringdistillation.
2L PremiumShort Path Distillation Kit By BVV
During this time, an idealvacuumpressure before filling and then poorvacuum performance after filling will likely cause an incompletedevolatilization. If this is the case, allow the oil to finishdevolatilization andvacuum performance to improve before proceeding withdistillation. Start by turning on the heated mantle setting the mantle temperature to 50C/122F and the magnetic stir RPM on a low until theviscosity of thebotanical extractsolution has decreased enough for mixing to occur. During thedistillation process, both heated mantle temperature and magnetic stir bar RPM can increase the rate ofevaporation. Keep in mind the rate ofevaporation should never exceed the capability of the condenser or cold trap. For this reason a steady increase of heat should be used and stir bar RPM should be optimized to create ideal turbulence of the liquid and maintained during the entirety of the run.
28mm L-Style MagneticPTFE Stirrer By BVV
Once the heated mantle reaches the set temperature of 50C/122F, proceed to increase mantle temperature in 20C/68F increments as the mantle operating temperature reaches the set temperature point until the extract is observed to boil. Once boiling is observed, mantle temperature can now be increased based on an observed drop indistillation headvapor temperature or using a decreased rate of boiling or reflux as indicators to increase mantle temperature. For the latter, once a 1-degree drop indistillation headvapor temperature is observed, proceed to increase mantle temperature in 5C/41F degree increments until the time comes to swap collections flasks and collect a different fraction.
2L Digital heating and Stirring Mantle By BVV
It is highly recommended to increase slowly and steadily as to prevent bumping over or over-reacting inside the boiling flask.Short pathdistillation can be slow to start, and the operator should maintain presence and monitor the system closely as temperatures are increasing. Keep in mind that during this time, asdistillation headvapor temperature increases, condenser temperature can also be increased to maintain a 20C colder condenser than thedistillation headvapor temperature, or it can be kept at the same temperature for the entirety of thedistillation procedure, typically between a range of 70-90C/158-194F.
Asdistillation continues, whendistillateviscosityis increased, or an increased flow ofdistillate is observed, both can be a good indicator that the target compounds are starting to distill, and the collection flask should be changed to isolate the main body from the heads fraction.
To ensure the main body fraction is of the highest purity, it is common to allow some of the main body to collect into the heads fraction before swapping flasks; this ensures that the lighter volatile compounds are not collected into the main body fraction. Once the main body is assumed to be distilling prior to swapping the flask, it's best to allow evaporation at that temperature range to diminish, observed by a significant drop in head vapor temperature and a visual reduction of boiling to ensure the lower boiling point compounds have distilled prior to swapping collection flasks and increasing mantle temperature to evaporate the target compounds of the main body fraction at a higher temperature range.
ValvedVacuum Adapter 2.0 By BVV
When it comes time to swap flasks and separate fractions, this process can differ depending on the collection apparatus. When using an isolation valve, start by turning the isolation valve clockwise to isolate the receiving flask, then open the bleeder valve to break thevacuum on the receiving flask before proceeding to change flasks. Once the flasks have been changed first seal the bleeder valve before slowly turning the isolation valve counterclockwise to connect the collection flask back to the system and proceed toevaporate and collect the main body fraction.
As thedistillation process is resumed and theevaporation of the main body is achieved. Continue to ramp mantle temperature in 5C increments as a 1-degree drop in headvapor temperature is observed. Depending on personal preference, once satisfied with the collection of the main body,distillation can be stopped leaving the tails fraction remaining in the resinous remains of the starting material or the main body flask can be changed yet again to capture the tails fraction of thedistillation. Typically a visual darkening of condensate is used as an indication to swap flask yet again to collect the tails fraction in a separate fraction. While the tails fraction can contain some desired compounds, these are typically separated from the main body to achieve a main body of the highest purity and preserve a lighter colored main body.
Once satisfied with the collection of either the main body of the tails fraction, proceed to end thedistillation run by first reducing the mantle temperature set point to 100C/212F. As the mantle temperature falls below 150C/302F, proceed to close thevacuumvalve, turn thevacuum pump off, and slowly releasevacuum from the system allowing system pressure to equalize. The collection flask can now be disassembled and transferred to a storage vessel. Before transferring thedistillate, ensure thevacuum grease is cleaned from the collection flask joints before transferring thedistillate to another vessel. Now that thedistillate has been collected it can be redistilled which is referred to as a second pass to further refine the main body fraction resulting in an even purer collection of target compounds or the first passdistillate can be used for further refinement via isolation.
SHORT-PATH DISTILLATION STANDARD OPERATING PROCEDURE
Purpose
The purpose of this procedure is to provide detailed instructions for short pathdistillation of abotanical extract.
Scope
This procedure applies to all lab technicians tasked withbotanical extractdistillation.
Definitions/Acronyms
Personal Protection Equipment (PPE):Items worn to protect employees from exposure to hazardous materials and prevention of injury.
Safety Data Sheet (SDS):Provides useful information on chemicals, describing the hazards the chemical presents, and giving information on handling, storage, and emergency measures in case of an accident.
Distillation:is The action of purifying a liquid by a process of heating and cooling.
Decarboxylation: is a chemical reaction that removes a carboxyl group from an organic compound resulting in the release of carbon dioxide and the conversion of a compound into a more distillable form.
Vacuum:avacuum is created by removing air from a container using avacuum pump evacuating the space entirely of matter.
Micron/mTorr: are interchangeable units of measure forvacuum depth represented as one-millionth of a meter of mercury displacement undervacuum.
Safety
SDS Sheets: No additional chemicals are used in this process.
PPE: The following should be worn by all lab personnel while performingbotanical extractdistillation:
Protective eyewear
Lab coat
Gloves
5.Hazard Identification
Preparation and Use:
A botanical extractwill be heated undervacuum to distill target compounds
Concentration- Winterized and decarboxylatedbotanical extract.
Quantity- >60% boiling flask volume.
Frequency- An initial volume of thebotanical extractis used, no more are added.
Location-Distillation occurs inside ashort path distillationstill undervacuum within a well-ventilated area.
Potential Hazards and Risks
The explosion from excessive internal pressure: theevaporator flask and/or the condenser could explode if the internal pressure produced byevaporation becomes too great.
Burns: Do not touch the heated mantle orevaporating flask during operation.
Brokenglassware: Cracks or chips inglassware can result in glass rupturing causing chemical exposure and explosion hazard to the user. Always inspectglassware prior to use.
6. Preparation
Assembly
Stage theshort path distillation apparatus in a well-ventilated area, where thevacuum pump exhaust can be vented outside the facility.
Assemble theshort pathdistillation system, ensuring each joining is properly greased by applying a small amount ofvacuum greaseon the upper half of the male joint and inserting it into the female joint. Once in place, rotate the male joint 360 degrees until an even coating ofvacuum grease can be seen across the entire circumference of the upper half of the male joint. (Fordistillations at temperatures above 204C/399F, useApizon High-TemperatureVacuum Grease.)
When a complete seal ofvacuum grease has been made, secure the joint with an appropriately sizedkeck clipand support the component with the appropriate lab stand support or scissor jack.
Ensure all process connections are appropriately sealed, and allglassware is appropriately supported by the appropriate lab stands and scissor jacks and proceed tovacuum test the system.
Ensure thevacuum pump is appropriately filled with freshvacuum pump oil and close thevacuum valve to the system before turning on thevacuum pump in opengas ballastmode and allowing it to warm up for 30 minutes to an hour.
Once thevacuum pump oil has been appropriately warmed up, proceed to open thevacuum valve allowing the entire system to be pulled undervacuum.
At this point, monitor thevacuum gauge, if deepvacuum levels are not attained, proceed to identify thevacuum leak by spraying each glass joint with soapy water, turning off thevacuum pump, and bleeding pressure before disassembling and reassembling connections that are not maintaining a seal.
Proceed tovacuum test and disassemble and reassemble the system as needed until there are novacuum leaks andvacuum levels can be maintained.
After thevacuum testing is complete, proceed to connect the heated circulator to the condenser port and fill the heated circulator with the appropriate heat transfer fluid. If operating the heated circulator in temperature ranges below 90C/194F, distilled water or a water-glycol mixture is sufficient. If operating a heated circulator above 90C/194F, a high-temperature heat transfer fluid like silicone oil is recommended. When connecting the heated circulator to the condenser, ensure the heat transfer fluid inlet is flowing from the top of the condenser to the bottom and all clamps are tightened appropriately.
Preparation
To prepare forshort pathdistillation, start by allowing thevacuum pump to warm up in open ballast mode and bringing the system components to operating temperature.
Ensure thevacuumpump oil is filled to the appropriate operating level and thevacuum valve to the system is closed. Proceed to turn on thevacuum pump in open ballast mode and allow it to warm up for 30 minutes to an hour beforedistillationwhile the rest of the system is prepared.
While thevacuum pump is warming up in open ballast mode, proceed to bring the cold trap to operating temperature by turning on the cold trap immersion probe chiller setting it to its lowest setting, or by preparing the cold trap with a dry ice/ethanol slurry by filling the cold trap 1/2 way withethanol and slowly adding dry ice to avoid over bubbling of thesolution.
As the cold trap reaches operating temperature, proceed to bring the condenser's heated circulator to operating temperature by first ensuring that the circulator bath is filled to the appropriate fill level and the fluid circulation is connected to flow into the top of the condenser and out of the bottom of the condenser.
Proceed to ensure all clamps are tightened appropriately before turning on the system and setting it to the desired condenser temperature. For the start of the run, a temperature of 40C/104F can be used, and this incrementally increased, asvapor temperature rises to maintain a 20C degree difference betweenvapor temperature and condenser temperature, or the condenser can be set to a temperature range of 70-90C/158-194F for the entirety of the run.
As thevacuum pump warms up and the system reaches the desired operating temperature, proceed to load thebotanical extractinto the boiling flask by removing the mantle thermal probe, cleaning the joint of anyvacuum grease, and attaching a glass funnel. Proceed to fill the boiling flask with winterized and devolitlizedbotanical extract. The boiling flask should be filled to a minimum of 25% of the capacity of the boiling flask, ensuring the there is enoughsolution for the thermal probe to read, and no more than 50% of the total volume of the boiling flask to ensure the there is enough headspace for thesolution to boil. Keep in mind winterized and devolitilizedbotanical extractis ideal forshort pathdistillation as unwinterizedcrude oilwill make cleaning the boiling flask a pain, and non-devolitized crude may impede its ability to reach lowvacuum levels.
Once thebotanical extracthas been loaded into the boiling flask, add the magnetic stir bar and reinstall the mantle temperature probe into the boiling flask ensuring the probe reaches 1/4" from the bottom of the boiling flask without touching the bottom of the flask or interfering with the rotation of the magnetic stir bar. The system is now sealed and can bevacuum tested once again to ensure a deepvacuum is reached before applying heat to the system.
Once thevacuum pump has operated in closed ballast mode for a minimum of 30 minutes, proceed to switch thevacuum pump into closed gas ballast mode and slowly open thevacuum valve to pull the system under fullvacuum and start theshort path distillation procedure.
6. Procedure
Turn on the heated mantle and apply heat to begin the process ofdistillation. Start by setting the mantle temperature to 50C/122F and the magnetic stir RPM on a low until theviscosity of the botanical extractsolution has decreased enough for mixing to occur. During thedistillation process, both heated mantle temperature and magnetic stir bar RPM can be used to increase the rate ofevaporation. Keep in mind the rate ofevaporation should never exceed the capability of the condenser or cold trap. For this reason, a steady increase of heat should be utilized, and stir bar RPM should be optimized to create ideal turbulence of the liquid and maintained during the entirety of the run.
Once the heated mantle reaches the set temperature of 50C/122F, proceed to increase mantle temperature in 20C/68F increments as the mantle operating temperature reaches the set temperature point until the extract is observed to boil.
Once boiling is observed, mantle temperature can now be increased based on an observed drop indistillation headvapor temperature or using a decreased rate of boiling or reflux as indicators to increase mantle temperature. For the latter as a 1C or a 5F degree drop indistillation headvapor temperature is observed, proceed to increase mantle temperature in 5C/41F degree increments until the time comes to swap collections flasks and collect a different fraction.
During this time, asdistillation headvapor temperature increases, condenser temperature can also be increased to maintain a 20C colder condenser than thedistillation headvapor temperature, or it can be kept at the same temperature for the entirety of thedistillation procedure, typically between a range of 70-90C/158-194F.
Whendistillate viscosity is increased, or an increased flow ofdistillate is observed, both can be a good indicator that the target compounds are starting to distill, and the collection flask should be changed to isolate the main body from the heads fraction.
To ensure the main body fraction is of the highest purity it allows some of the main body to collect into the heads fraction before swapping flasks; this ensures that the lighter volatile compounds are not collected into the main body fraction.
Once the main body is assumed to be distilling prior to swapping the flask, allowevaporation at that temperature range to diminish, observed by a significant drop in headvapor temperature and a visual reduction of boiling to ensure the lowerboiling pointcompounds have distilled prior to swapping collection flasks and increasing mantle temperature toevaporate the target compounds of the main body fraction at a higher temperature range.
When it comes time to swap flasks and separate fractions, this process can differ depending on the collection apparatus. When using an isolation valve, start by turning the isolation valve clockwise to isolate the receiving flask. then open the bleeder valve to break thevacuum on the receiving flask before proceeding to change flasks. Once the flasks have been changed first seal the bleeder valve before slowly turning the isolation valve counterclockwise to connect the collection flask back to the system and proceed toevaporate and collect the main body fraction.
As thedistillation process is resumed and theevaporation of the main body is achieved, continue to ramp mantle temperature in 5C increments as a 1-degree drop in headvapor temperature is observed. Depending on personal preference, once satisfied with the collection of the main body,distillation can be stopped leaving the tails fraction remaining in the resinous remains of the starting material, or the main body flask can be changed yet again to capture the tails fraction of thedistillation. Typically, a visible darkening of condensate is used as an indication to swap flasks and collect the tails fraction in a separate fraction. While the tails fraction can contain some desired compounds, these are typically separated from the main body to achieve a main body of the highest purity and preserve a lighter-colored main body. Keep in mind thatdistillation should not be continued past a mantle temperature of 220C/428F.
Once satisfied with the collection of either the main body or tails fraction, proceed to end thedistillation run by first reducing the mantle temperature set point to 100C/212F.
As the mantle temperature falls below 150C/302F, proceed to close thevacuum valve, turn thevacuum pump off, and slowly release thevacuum from the system allowing system pressure to equalize.
The collection flask can now be disassembled and transferred to a storage vessel. Before transferring thedistillate, ensure thevacuum grease is cleaned from the collection flask joints before transferring thedistillate to another vessel.
The refineddistillate can now be redistilled to refine the main body fraction resulting in an even purer collection of target compounds or be used for further isolation of target compounds via isolation.